All Schrödinger offices worldwide will be closed for the week of August 17-21 as part of a company-wide initiative to rest and recharge. Please expect limited responses during this time. Scientific and Technical Support team members will be available to answer emergency support issues only.

Festival of Biologics 2025

Conference

Festival of Biologics 2025

CalendarDate & Time
  • April 23rd-24th, 2025
LocationLocation
  • San Diego, California

Schrödinger is excited to be participating in the Festival of Biologics 2025 conference taking place on April 23rd – 24th in San Diego, California. Join us for a poster presentation by Zhe Mei, Senior Scientist I at Schrödinger, titled “Antibody Optimization with Physics based and Machine Learning based modeling.” Stop by booth #835 to speak with Schrödinger scientists.

icon time 6:10 PM
Antibody Optimization with Physics based and Machine Learning based modeling

Speaker:
Zhe Mei, Senior Scientist I, Schrödinger

Abstract:
Optimizing antibody properties, such as binding affinity, stability and aggregation is crucial for developing safe and effective biotherapeutics. This work presents an integrated approach leveraging physics-based modeling and machine learning to address these challenges. We use both methods to predict 3D structures and calculate a rich set of sequence-based, structure-based and surface patch-based protein descriptors that can be used to train machine learning models. Further, we can identify hotspots for targeted optimization of stability and affinity and apply physics-based methods like free energy perturbation (FEP+) to design improved and developable variants.

MS Batteries – Michael Rauch

Molecular Insight, Material Impact

Molecular Insight, Material Impact

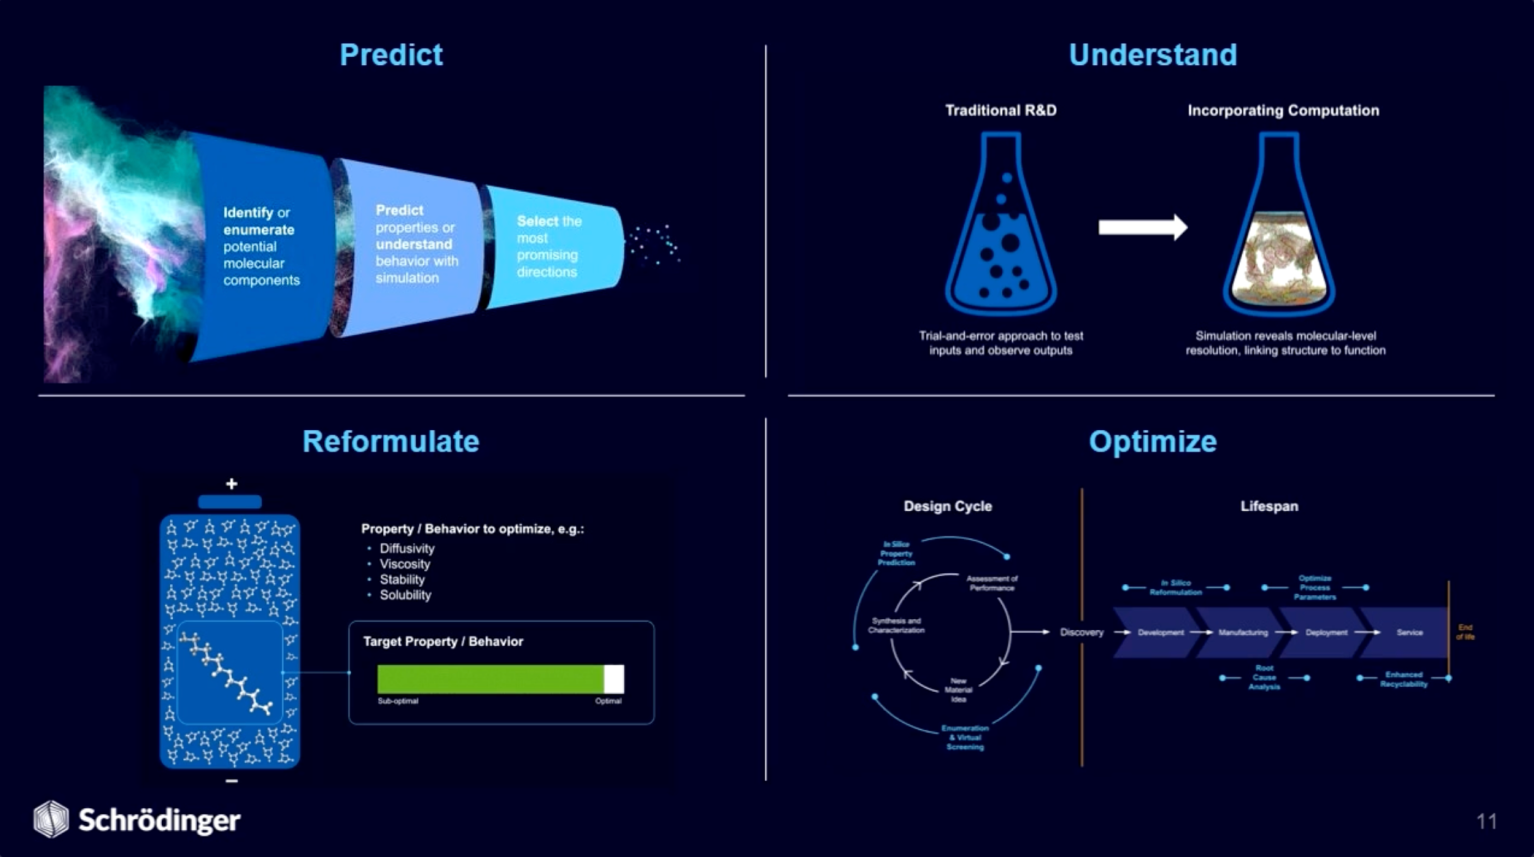
Please find within this page details around utilizing Schrödinger’s software and services within the Battery industry.

Battery Presentation

Download our Battery presentation to learn more about Schrödinger and our software capabilities.

Typical Roadmap to Adoption

Here is an example of an adoption pattern followed by similar companies. This is completely customizable and can be segmented to meet your needs.

1

Technical Engagement

Meet with our experts to discuss how modeling and simulation can address your R&D needs

2

Contract Research

Optionally, outsource a project to our experienced contract research team

3

Training & Courses

Learn to use our software – whether you are a complete beginner or expert

4

Software Adoption

Bring the tools that you need in-house to meet your R&D goals

5

Ongoing Support

Receive premium support from our expert scientific team

Webinars and White Papers

High-performance materials discovery: A decade of cloud-enabled breakthroughs Webinar Materials Science
High-performance materials discovery: A decade of cloud-enabled breakthroughs

This talk will showcase how Schrödinger’s integrated materials science platform enables massive parallel screening and de novo design campaigns across diverse applications.

How Physics-based Modeling and Machine Learning Enable Accelerated Development of Battery Materials Webinar Materials Science
How Physics-based Modeling and Machine Learning Enable Accelerated Development of Battery Materials

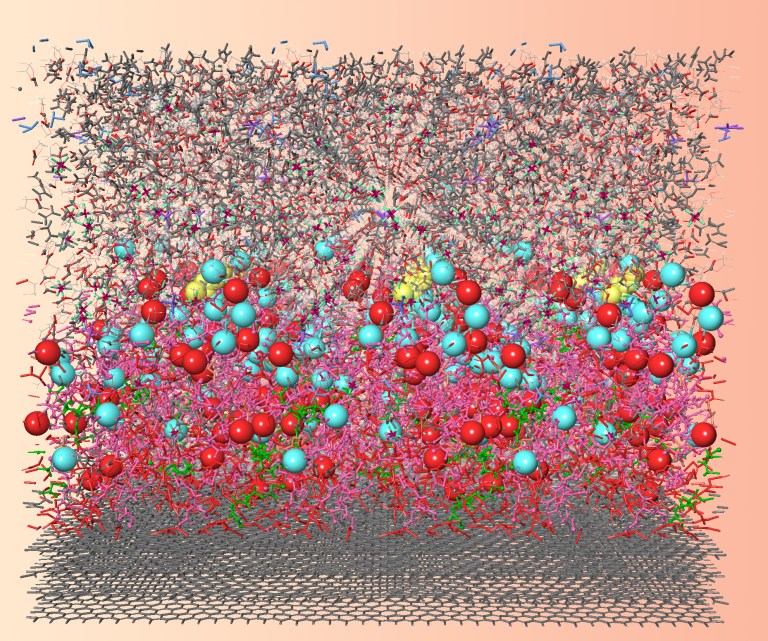
In this webinar, we focus on examples to demonstrate the application of automated solutions for accurate prediction of thermodynamic stability and voltage profile of cathode materials, ion diffusion pathways and kinetics in electrode materials, transport properties of liquid electrolytes and modeling the nucleation and growth of solid electrolyte interphase (SEI) layers using Schrödinger’s SEI simulator module.

Electrodes, electrolytes & interfaces: Harnessing molecular simulation and machine learning for rapid advancements in battery materials development Webinar Materials Science
Electrodes, electrolytes & interfaces: Harnessing molecular simulation and machine learning for rapid advancements in battery materials development

In this webinar, we demonstrate the application of automated solutions for accurate prediction of electrode materials.

Purposeful simulation: Maximising impact in surface chemistry modelling Webinar Materials Science
Purposeful simulation: Maximising impact in surface chemistry modelling

In this webinar, learn about a variety of atomistic models of surfaces and gain perspective on the underlying rationale, benefits and limitations of each.

Publications

Exploring Molecules with Low Viscosity: Using Physics-Based Simulations and De Novo Design by Applying Reinforcement Learning

Panasonic Publication

Read
Designing the next generation of polymers with machine learning and physics-based models

SABIC Publication

Read
Quantum chemical package Jaguar: A survey of recent developments and unique features

Schrödinger Publication

Read
Leveraging high-throughput molecular simulations and machine learning for the design of chemical mixtures

Schrödinger Publication

Read
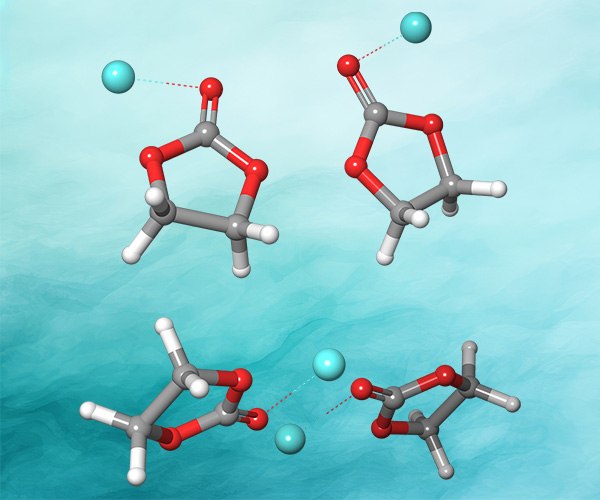
Accurate Quantum Chemical Reaction Energies for Lithium-Mediated Electrolyte Decomposition and Evaluation of Density Functional Approximations

Academic Publication

Read
High-Dimensional Neural Network Potential for Liquid Electrolyte Simulations

Schrödinger Publication

Read

Educator’s Month: Molecules & Models – A Virtual Science Fair

Virtual Science Fair

Educator’s Month – Molecules & models: A virtual science fair

CalendarDate & Time
  • June 12th, 2025
LocationLocation
  • Virtual

As part of Educator’s Month, Schrödinger hosted its first Virtual Science Fair on June 12, 2025. This free event invited undergraduate students from across the U.S. to showcase their research, engage in discussions with Schrödinger judges, and compete for awards recognizing their creativity, effort, and commitment.

The Virtual Science Fair was open to first-time undergraduate participants from a wide range of disciplines. All projects were required to include a computational component, such as artificial intelligence, experimental design, or molecular modeling in fields including drug discovery, agrochemicals, materials science, medicinal and organic chemistry, pharmaceuticals, polymers, catalysis, computational biology, biophysics, or theoretical chemistry.

Winners received a cash prize and one year of unlimited access to Schrödinger’s online certification courses – supporting both their research and ongoing computational skill development.

Presentation Recordings

  • Computational Design of de novo Transcription Factors for Targeted Genetic Repression
    Speaker:
    Beau Lonnquist, University of Washington, (science fair winner)
    Watch now
  • Modeling Molecular Scale Dynamics of Kinetically Gated Carbon Dioxide Capture Using Photoswitch Functionality in Metal Organic Frameworks
    Speaker:
    Ryan Miller, Pacific University, (science fair winner)
    Watch now
  • Molecular Basis of Adenylyl Cyclase 1 Activation Revealed by MD Simulations
    Speaker:
    Shreya Krishnan, Purdue University, (science fair winner)
    Watch now
     
  • Repurposing L-Type Calcium Channel Blockers as Respiratory Virus Therapeutics: A Computational Modeling Approach
    Speaker:
    Aiden T. Day, Saint Joseph’s College of Maine
    Watch now
  • Discovery of FabG Inhibitors for Yersinia pestis Using Computational and Biochemical Approaches
    Speaker:
    Catalina Colling, University of Texas at Austin
    Watch now
     
  • Analyzing the Value of Machine Learning in Improving the Acceptance Rate for Metropolis Monte Carlos
    Speaker:
    Enoch Woldu, University of Chicago
    Watch now
  • Comparative Molecular Drug Docking to hERG and CaV1.2- Channels to Understand Drug-Induced Cardiac Risks
    Speaker:
    Ensley Jang, University of California, Davis
    Watch now
     
  • Computational Development of a Hydrolase with Increased Degradation Capabilities Against Crystalline PET
    Speaker:
    Mena Boggs, NCSSM/NC State University
    Watch now
  • Protein and Solvent Dynamics Simulations to Understand Cancer Mutations
    Speaker:
    Michael Sarullo, Yale University
    Watch now
     
  • Dynamic Docking: A Scalable Computational Framework for Conformational Profiling of Small Molecule/RNA Binding
    Speaker:
    Nakul Balaji, Florida Atlantic University
    Watch now
  • Discovery of Aza-stilbene as a Scaffold for a Histamine Receptor H2 Antagonist for the Treatment of Gastroesophageal Reflux Disease
    Speaker:
    Nihar Kummetha, North Carolina School of Science and Mathematics
    Watch now
     
  • Analyzing Quantum Exceptional Point Invisibility for Experimentally Realizable Triple-Gaussian Potentials
    Speaker:
    Shrikar Dulam, University of Illinois, Urbana-Champaign
    Watch now
  • Deep Learning–Based Structural Modeling of YscF Mutants Reveals Determinants of Type III Secretion System Architecture in Yersinia pestis
    Speaker:
    Stephanie Bellido, Nova Southeastern University
    Watch now
     
  • Extending the Functionality of the Excel-to-SBOL Converter for Broader Synthetic Biology Applications
    Speaker:
    Taisiia Sherstiukova, University of Colorado Boulder
    Watch now
  • BitBIRCH: Efficiently Clustering 1 Billion Molecules
    Speaker:
    Vicky Jung, University of Florida
    Watch now

SAMPE 2025

Conference

SAMPE 2025

CalendarDate & Time
  • May 19th-22nd, 2025
LocationLocation
  • Indianapolis, Indiana

Schrödinger is excited to be participating in the SAMPE 2025 conference taking place on May 19th – 22nd in Indianapolis, Indiana. Join us for a presentation by Andrea Browning, Director of Polymers and Soft Matter at Schrödinger, titled “Aiding Sustainable Composites Development with Simulation of Natural Fibers.” Stop by booth U15 to speak with Schrödinger scientists.

icon time MAY 20 | 10:30AM
Accelerating innovation in advanced composites with a digital chemistry platform

Speaker:
Andrea Browning, Director of Polymers and Soft Matter, Schrödinger

Abstract:
Demands for advanced materials and processes have grown as various industries search for improved and lower cost solutions. Their development has traditionally relied on experimental exploration of candidate chemistries, which is time-consuming, expensive and limited in scope. Optimizing key properties of advanced composites, coatings and energy storage requires understanding of their chemistry and microstructure at atomic level. Physics-based modeling and chemistry-informed machine learning (ML) can significantly accelerate the development process from material selection to processing and lifetime analysis, ensuring that target performance is met. In this presentation, we will show how Schrödinger’s digital chemistry technology can catalyze formulation development through efficient screening of candidate mixtures and increase fundamental understanding of structure-property relationships. We will showcase how our AI/ML and physics-based tools can be applied to the screening of polymer chemistries for target thermomechanical properties for thermosets, efficient additive selection for complex coatings and reactivity at interfaces.

icon time MAY 21 | 3:00PM
icon location Room 126
Aiding Sustainable Composites Development with Simulation of Natural Fibers

Speaker:
Andrea Browning, Director of Polymers and Soft Matter, Schrödinger

Abstract:
Natural fibers have gained interest as potential components to improve the overall sustainability of composite materials. However, natural fibers have unique challenges and cannot be simply substituted for carbon or glass fibers. Better understanding of how natural fibers behave with standard and new resins, along with how they can degrade can help to reduce the risk in transitioning to these new materials. Molecular scale simulation is a powerful tool to provide that understanding. The interaction between resin and natural fibers as well as the impact of compatibilizers are important in designing a composite formulation for use with natural fibers. The degradation of natural fiber chemistry, cellulose, is also insightful as part of the recycling potential for natural fiber composites. This study will present molecular level simulations that address both the natural fiber composite mechanical properties, as well as degradation of cellulose. These findings highlight the impact of molecular simulations to bridge the connection between molecular level interactions and design considerations in natural fiber composites, aiding in the design of more sustainable composites.

Accelerating OLED innovation with multi-scale, multi-physics simulations

APRIL 16, 2025

Accelerating OLED innovation with multi-scale, multi-physics simulations

OLED technology is widely used in mobile devices, AR/VR systems, and automotive displays, with its flexibility enabling foldable devices and enhanced user interactions. However, advancing OLEDs requires overcoming challenges such as improving efficiency, extending lifetime, ensuring color stability, and optimizing scalable manufacturing. Since device performance depends on both material properties and fabrication methods, a deeper understanding of OLED materials and device architectures is essential for innovation.

Traditional trial-and-error approaches to materials discovery are costly and time-consuming. To address this, we present the synergistic application of Ansys and Schrödinger predictive technologies to accelerate OLED development through a multi-scale, multi-physics simulation approach. This framework integrates:

  • Molecular modeling to predict materials properties at atomistic scale
  • Nanoscale simulations to examine photonic responses and light-matter interactions
  • Macroscale modeling to assess human perception of displays in real-world conditions

By combining Schrödinger’s expertise in molecular simulations with Ansys’s advanced device modeling, this approach enables faster, cost-effective OLED innovation. Join us to explore how integrated digital workflows drive the design of next-generation, high-performance OLEDs.

Our Speakers

Hadi Abroshan

Principal Scientist I, Schrödinger

Hadi Abroshan is the Product Manager for Organic Electronics at Schrödinger. He holds a Ph.D. from Carnegie Mellon University and has conducted research at Stanford University and Georgia Tech. Hadi specializes in multiscale simulations, leading projects to design cost-effective multifunctional materials for optoelectronics. His expertise lies in developing computational strategies that bridge atomistic structures to multilayered device scales, using a blend of physics-based methodologies and machine learning techniques. His work has led to the discovery of novel, environmentally friendly materials and processes with superior efficiencies.

Thibault Leportier

Senior R&D Engineer, Ansys

Thibault Leportier is an optical scientist with expertise in geometrical and Fourier optics, lasers, and metrology. Holding a Ph.D. in 3D display technologies and digital holography, Thibault has worked extensively on waveguide design for smart glasses and eye-tracking applications. Currently, his focus is on developing innovative solutions to bridge nanoscale simulations with macroscale optical systems, including advancements in metalens and grating models.

Target enablement, preparation, & validation

Target Enablement Course Image of a ligand and protein

Target enablement, preparation, & validation


Enabling protein structures from x-ray crystallography, cryo-EM, ML-methods, and homology modeling for structure-based computational workflows

Details
Available Languages
Chinese, English, Japanese, Korean
Duration
Up to 20 hours over 5 weeks from selected start date
Level
Intermediate
Cost
$715 for non-student users
$270 for student / post-doc
Who should take this course?
Medicinal chemists, cheminformaticians, ML scientists, new computational chemists, and structural biologists

Overview

With the explosion of available structures from x-ray crystallography, cryo-EM and, more recently, machine learning (ML) methods, there is a growing need for tools and workflows for preparing, refining and validating structures for structure-based computational workflows.

Schrödinger’s online course, Target Enablement, Preparation, and Validation will provide expert guidance and best practices to equip you to enable projects for prospective structure-based computational modeling workflows such as virtual screening, molecular dynamics simulations, and free energy perturbation calculations.

This course is ideal for those who wish to develop professionally and expand their CV by earning certification and a badge.

  • Work hands-on with Schrödinger’s industry-leading Maestro and command line interface
  • Jump start your research program by learning methods that can be directly applied to ongoing projects
  • Learn topics ranging from refining AlphaFold structures to cryptic pocket identification
  • Independently perform a case study to demonstrate mastery of the course content
  • Benefit from review and feedback from Schrödinger Education Team experts for course assignments and course-related queries
  • Work on the course materials on your own schedule whenever convenient for you within the course session


This course comes with access to a web-based version of Schrödinger software with the necessary licenses and compute resources for the course:

Requirements
  • A computer with reliable high speed internet access (8 Mbps or better)
  • A mouse and/or external monitor (recommended but not required)
  • Working knowledge of general chemistry and structural biology
  • Working knowledge of Maestro. This course will not teach you how to navigate the Schrödinger graphical user interface, Maestro. Please work through our Getting Started with Maestro resources to become familiar with using Maestro.
Certification
  • A certificate signed by the Schrödinger course lead to add to your CV or resume
  • A badge that can be posted to social media, such as LinkedIn
background pattern

What you will learn

X-ray and cryo-EM structures

Learn best practices for preparing and refining experimental structures of varying quality

ML-predicted structures and homology modeling

Learn best practices for working with and refining ML-predicted structures (such as from AlphaFold) and homology models

Binding site identification

Learn how to evaluate the drugability of small molecule binding site, as well as search for and characterize potential cryptic pockets

Prospective enablement of a target

Apply your skills by independently enabling a target through thorough inspection of available structures, analysis, and refinement

Modules

Module 1
2 Hours

Target enablement methods and the value of structural validation

Video
Video

Course overview

Checkpoint
Syllabus and honor code

Expectations surrounding academic integrity

Video Tutorial
Videos
  • The importance of structure validation in computational experiments for drug discovery
  • Comparing common target enablement methods
End checkpoint
End of module checkpoint
Module 2
5 Hours

Starting point: X-ray crystal and cryogenic electron microscopy structures

Video
Video

Structure availability and experimental considerations

Tutorial
Tutorials:
  • Structure quality metrics
  • Inspection workflows
  • Protein preparation
  • Basic refinement
End checkpoint
End of module checkpoint
Module 3
4 Hours

Starting point: AlphaFold structures and homology models

Video
Video

Generating, inspecting, and validating AlphaFold and homology models

Tutorial
Tutorials
  • Obtaining and reviewing AlphaFold structures
  • Homology modeling
  • Model refinement methods
End checkpoint
End of module checkpoint
Module 4
5 Hours

Next steps: advanced preparation, refinement, and validation of structures

Video
Video

Refinement and validation methods for more challenging targets

Tutorial
Tutorials
  • Manual protein preparation
  • SiteMap
  • Mixed Solvent Molecular Dynamics
  • WaterMap
End checkpoint
End of module checkpoint
Module 5
4 Hours

Case study: prepare and validate a structure for computational drug discovery

Video
Video
  • Case study overview
  • Case study findings and course closing
Tutorial
Tutorial

Structural inspection, preparation, and validation

Assignment
Assignment

Review and discuss case study findings

Course completion
Course completion and certification

Need help obtaining funding for a Schrödinger Online Course?

We proudly support the next generation of scientists and are committed to providing opportunities to those with limited resources. Learn about your funding options for our online certification courses as a student, post-doc, or industry scientist and enroll today!

What our alumni say

“The course was designed to tackle the pressing need of drug discovery acceleration when high precision protein prediction methods are readily available. I highly recommend this course.”
Wei WangAssistant Professor, Icahn School of Medicine

Show off your newly acquired skills with a course badge and certificate

When you complete a course with us in molecular modeling and are ready to share what you learned with your colleagues and employers, you can share your certificate and badge on your LinkedIn profile.

Frequently asked questions

How much does the Target enablement, preparation, and validation online course cost?

Pricing varies by each course and by the participant type. For students wishing to take this, we offer a student price of $240, and $645 for non-students.

What time are the lectures?

Once the course session begins, all lectures are asynchronous and you can view the self-paced videos, tutorials, and assignments at your convenience.

How could I pay for this course?

Interested participants can pay for the course by completing their registration and using the credit card portal for an instant sign up. Please note that a credit card is required as we do not accept debit cards. Additionally, we can provide a purchase order upon request, please email online-learning@schrodinger.com if you are interested in this option. If you have any questions regarding how to pay for the course, please visit our funding options page.

Are there any scholarship opportunities available for students?

Schrödinger is committed to supporting students with limited resources. Schrödinger’s mission is to improve human health and quality of life by transforming the way therapeutics and materials are discovered. Schrödinger proudly supports the next generation of scientists. We have created a scholarship program that is open to full-time students or post-docs to students who can demonstrate financial need, and have a statement of support from the academic advisor. Please complete the application form if you qualify for our scholarship program!

Will material still be available after a course ends?

While access to the software will end when the course closes, some of the material within the course (slides, papers, and tutorials) are available for download so that you can refer back to it after the course. Other materials, such as videos, quizzes, and access to the software, will only be available for the duration of the course.

Do I need access to the software to be able to do the course? Do I have to purchase the software separately?

For the duration of the course, you will have access to a web-based version of Maestro, Bioluminate, Materials Science Maestro and/or LiveDesign (depending on the course). You do not have to separately purchase access to any software. While access to the software will end when the course closes, some of the material within the course (slides, papers, and tutorials) are available for download so that you can refer back to it after the course. Other materials, such as videos, quizzes, and access to the software, will only be available for the duration of the course. Please note that Schrödinger software is only to be used for course-related purposes.

Related courses

Designing Quality Ligand Libraries_Featured image Life Science Life Science
Designing quality ligand libraries

Exploring chemical space, profiling and tailoring ligand libraries, validating docking models, and methods of enumeration for hit discovery

Free Energy Calculation for Drug Design with FEP+ Life Science Life Science
Free energy calculations for drug design with FEP+

Running, analyzing, and troubleshooting relative binding FEP+ calculations for small molecule lead optimization

ACS Spring 2025

ACS Spring 2025

CalendarDate & Time
  • March 23rd-27th, 2025
LocationLocation
  • San Diego, California

Schrödinger is excited to be participating in the ACS Spring 2025 conference taking place on March 23rd – 27th in San Diego California. Stop by booth #3532 to speak with Schrödinger scientists or join us for happy hour on March 26th. Full event details below.

icon time MAR 23 | 8:00AM
icon location Ballroom 20D
Unlocking a New Era: How AI is Transforming Drug Discovery and Development

Speaker:
Yefen Zou, Senior Principal Scientist, Schrödinger

Abstract:
We are seeing an unprecedented growth on AI-driven research towards the development of new medicines. Nowadays, AI can impact all aspects of drug development: from enabling generative chemistry, to analysis of large datasets to accelerate hit finding, reduce timelines in the design of drug-like lead compounds, to structure-based drug design with AlphaFold to applications to green and sustainable drug development synthesis. This exponential growth covers AI-biotechs to large pharma, pioneer academic researchers and AI-technology organizations, effectively unlocking a new era to bring new medicines to patients in need. This symposium will showcase examples of how AI is impacting drug development from early discovery, to drug development, from industry to academic research and teams comprising members of MEDI, ORGN, COMP, I&EC, CINF Divisions.

icon time MAR 23 | 3:45PM
icon location Pacific Ballroom: Section 23
Transforming Polymer Design for Industrial Applications Using Experiment Data, Machine Learning, and Physics-Based Modeling

Speaker:
Atif Afzal, Principal Scientist II, Schrödinger

Abstract:
Designing industrially relevant polymers is challenging due to the need to simultaneously optimize multiple material properties, making traditional trial-and-error methods costly and inefficient. A promising alternative is the integration of machine learning (ML) and physics-based approaches to explore the polymer design space and identify candidates that meet specific industrial criteria. This work presents a workflow combining ML and molecular modeling techniques, demonstrated through two case studies in polymer design. The first case is an internal study focused on elastomers with target thermophysical and mechanical properties. Using curated datasets of experimental property values from the literature, we developed ML models that accurately predict the properties of new elastomers. These models screened over 100,000 copolymers, identifying top candidates with superior glass transition temperatures and elastic moduli suitable for elastomer applications. The second case presents results from a collaboration with SABIC Specialties involving polycarbonate design*, targeting key polymer properties, including glass transition temperature, optical properties, and mechanical strength. Using an experimental dataset, we trained ML models to predict these polymer properties based on structure and monomer composition. We subsequently screened several thousands of polycarbonate structures generated via R-group enumeration and identified the top candidates. These candidates were validated through molecular dynamics simulations and density functional theory, demonstrating strong correlations between ML predictions and physics-based approaches. Experimental validation further confirmed the accuracy of our models. This workflow demonstrates the power of integrating data-driven and physics-based methods in polymer design, offering an efficient strategy for material scientists working with limited experimental data. *Collaboration with SABIC Specialties is gratefully acknowledged.

icon time MAR 24 | 11:35AM
Combined Physics-Based and Machine Learning Approaches in the Design of Complex Materials

Speaker:
Anand Chandrasekaran, Senior Principal Scientist, Schrödinger

Abstract:
The simulation of materials properties using physics-based approaches, such as density functional theory (DFT) and molecular dynamics (MD), has long been successful in providing insights into structure-property relationships and subsequently aiding in the design of novel materials. More recently, machine learning (ML) has been used extensively in conjunction with physics-based modeling techniques to greatly accelerate materials innovation. The accuracy and generalizability of physics-based modeling improves the performance of ML models and enables them to be used effectively even in small-data regimes. Conversely, the speed and flexibility of ML help bridge the time- and spatial- scale limitations of physics-based models, creating a synergistic approach that optimizes both predictive accuracy and computational efficiency. In this talk, we demonstrate the application of this combined approach in designing materials and formulations across diverse applications, from battery electrolytes and fuel mixtures to thermoplastics and OLED devices. For instance, we demonstrate how DFT descriptors greatly improve the accuracy of ML models for optoelectronic molecules and battery electrolytes while descriptors from MD simulations can lead to better models for viscosity of organic molecules. We use Schrodinger’s automated Formulation ML solution, which takes into account both chemistry and composition, to train ML models for solubility of APIs in binary solvents and for the prediction of motor octane number of hydrocarbons. Additionally, we showcase recent advancements in our machine learning force field technology (MPQRNN), which has been trained on a vast chemical space encompassing over 86 elements, and demonstrate its application in accurately modeling the bulk properties of inorganic cathode coating materials.

icon time MAR 25 | 10:30AM
icon location Room 25C
Leveraging Cloud Computing to Efficiently Identify the Most Promising Compounds in Ultra-Large Chemical Spaces for In-Silico Hit Discovery

Speaker:
Steven Jerome, Executive Director, Schrödinger

Abstract:
By screening ultra-large libraries in the cloud with a per-target tailored, hierarchical screening approach, the dream of achieving double-digit hit rates and diverse starting chemical matter in virtual screening has been achieved for multiple drug discovery projects. Virtual libraries for in-silico hit finding can be thought of as curated subsets of larger chemical spaces defined by a set of reactions and matching reagent libraries. While many commercial vendors provide such “ready to screen” virtual libraries, advances in cloud computing make it possible to tailor custom libraries on a per-project basis by exploring the full vendor space. At Schrödinger, project teams begin all virtual screening campaigns by searching an internal cloud database built on Google’s BigQuery comprising more than 145 billion purchasable compounds representing 50 vendors, including fully enumerated ultra-large vendor spaces such as Enamine Real. In order to identify the most promising compounds for virtual screening, we have developed a screening methodology called QuickShape based on a custom 1D fingerprint which aims to capture pharmacophore-like features. This compact representation is well-suited for representing ultra-large chemical spaces and is incorporated into our cloud database. In this talk, we present our cloud-native approach for the generation of target-specific libraries for virtual screening together with a pair of prospective studies inside active drug discovery programs, covering both fragment and druglike molecule virtual screening.

icon time MAR 25 | 12:00PM
icon location Room 1A
20+ years of AI in Drug Discovery: From Promise to Impact

Panelist:
H. Rachel Lagiakos, Director, Medicinal Chemistry, Schrödinger

Abstract:
Talk followed by a panel discussion.

icon time MAR 26 | 9:00AM
icon location Room 28A/B
MEDI First Time Disclosures

Host:
H. Rachel Lagiakos, Director, Medicinal Chemistry, Schrödinger

Abstract:
The highly anticipated session that chronicles the journey of a molecule from its discovery on the bench to its progression into clinical trials. This session emphasizes the challenges and successes that medicinal chemists face every day, and reminds us of the tremendous impact our efforts can bring!

icon time MAR 26 | 6:00PM
MEDI First Time Disclosures Networking Reception

Hosted by Schrödinger
Join other medicinal chemists at this happy hour event for networking and drinks. Hosted outside room 28A/B in the San Diego Convention Center

Pharmaceutical Formulations Workshop 2025

Workshop

Pharmaceutical Formulations Workshop 2025

CalendarDate & Time
  • March 19th, 2025
  • 10:00AM CET
LocationLocation
  • Mannheim, Germany
Register

Using Schrödinger’s Materials Science Suite for molecular modeling and machine learning in the area of pharmaceutical formulation and delivery

Schrödinger invites you to a one-day in-person workshop in Mannheim, Germany to gain hands-on training in the use of our Materials Science Suite for drug development.

Participants will get practical experience and in-person guidance in using our Materials Science Suite and the tools involved in building molecules, polymers, and complex mixtures for use in molecular dynamics simulations. Leveraging automated property prediction workflows as well as analysis tools will play an important role. Another aspect will be the application of machine learning.

Examples of how molecular-scale simulations can inform drug delivery and formulation research will be included through-out the day.

When & Where:

Wednesday 19th March 2025, 10:00AM CET
Glücksteinallee 25
68163 Mannheim, Germany
(5 minutes walk from Mannheim Hauptbahnhof)

Please see the Agenda for more details about the content presented (Full agenda TBA).
Please see our FAQs for information regarding what to bring, getting to the venue, and accessibility.
If you need further information please contact Patrick Heasman: patrick.heasman@schrodinger.com
If you are interested but unable to attend in person, please reach out to the contact above.

Registration:

Registration is free and includes lunch and refreshments.

Participants must bring their own laptop to access the software, and an external mouse is recommended. We will be utilising our Virtual Computer, which is accessed via web browser – No software installation is required prior to the session.

Places are limited, so please ensure to register as soon as possible.

Registration will close at latest on Friday 14th March 2025

Who should attend:

Any researcher studying drug discovery, design, formulation, or interested generally in learning about computational materials science. No prior experience is required.

Instructional material can be reviewed before or after the workshop for free at:

https://www.schrodinger.com/sites/default/files/s3/release/current/Documentation/html/materials_science/tutorials_TOC.htm

Speakers and demonstrators:

Dr. Caroline Krauter
Dr. Irene Bechis
Dr. Patrick Heasman

Agenda

Register

FAQs

Where is the venue and how can I get there?

The workshop is being help at our offices in Mannheim, Germany. The building is accessible via car, and the train station is within a 5 minute walk.

What is included with my registration?

Registration is completely free to attend the workshop. We will also be providing food and refreshments throughout the day.

Can I join the session virtually / remotely?

As we want to give the attendees help and guidance during the workshop we currently have no intention of running this workshop online. Please reach out if you are interested but are unable to travel to the event location.

What do I need to bring?

A laptop is required for this workshop. We will not be providing any on the day, so please ensure that you bring one. We also recommend that you bring a personal laptop to avoid any firewall restrictions.

An external mouse is not required, but we do recommend that you bring one as our software makes full use of the 3 buttons.

No. We will be utilising the Schrödinger Virtual Computer for all hands-on sessions. A suitable web browser is required for accessing this (Chrome, Edge, Firefox).

How long is the workshop?

The workshop is an all day event to give you the best opportunity to learn about our tools and benefit from the practical sessions throughout. We will start at 10:00 am and finish at approximately 4:00 pm.

For accommodation and travel, we ask that attendees make their own arrangements. There are several hotels within walking distance to the venue, and the train station is situated close by.

Please contact Patrick Heasman (patrick.heasman@schrodinger.com) for any additional information about the event and the location.

Travel:

  • Via plane / train: Frankfurt / Frankfurt Airport – A direct train to Mannheim takes approximately 45 minutes.
  • Via car: There are several car parks located on Glücksteinallee.

Hotel recommendations:

  • Holiday Inn Mannheim City
  • LanzCarré Hotel Mannheim
  • Premier Inn Mannheim City Centre hotel
  • Hilton Garden Inn Mannheim

Release 2025-1

Library Background

Release Notes

Release 2025-1

Small Molecule Drug Discovery

Target Validation & Structure Enablement

Protein Preparation

  • Improved peptide bond connectivity by better integration of sequence information in the Protein Preparation Workflow (PPW)

Protein X-Ray Refinement

  • Phenix/OPLS can now run from CIF files containing reflections

Cryo-EM Model Refinement

  • Improved support of macrocycles in GlideEM/GlideXtal

Binding Site & Structure Analysis

Binding Site Characterization

  • Rationalize kinase selectivity challenges using the Kinase Conservation Analysis Interface that combines sequence and complex structural information: Identify promising selectivity handle residues to modulate ligand kinase selectivity that can be tested with FEP+ Residue Scan

Desmond Molecular Dynamics

  • Optimize the unbinding kinetics of protein-ligand complexes using dissolution rate predictions based on unbinding pathways identified by enhanced sampling methods

Mixed Solvent MD (MxMD)

  • Identify potential binding sites and assess drugability of competitive and allosteric binding sites with the full release of a Mixed Solvent Molecular Dynamics Interface to setup and analyze MxMD simulations

Hit Identification & Virtual Screening

Active Learning Applications

  • Improved diversity of top scoring ligands in Active Learning ABFEP by using 3D features extracted from Glide poses in ML model building
  • Researchers can now specify different batch sizes and selection rules to exploit or explore for each iteration in Active Learning simulations

Lead Optimization

Protein FEP

  • View trajectories and structural output from FEP+ Residue Scans in the FEP+ interface
  • Rationalize key interactions in ligand binding using per-contact residue interaction energy analysis which is new for ABFEP
  • Easily identify disconnected sub-maps in a busy FEP map

Spectroscopy

  • More accurate NMR spectra predictions by identifying magnetically inequivalent nuclei

Macrocycles

  • Automatically enumerate and cyclize peptide sequences from FASTA files with the new cyclize_peptide.py script
  • Ring template generation for Glide macrocycle docking with bespoke parameters is now automated by the macrocycle_template_gen.py script
  • An updated macrocycle_sample.py script with greater control of sampling options and a new receptor-aware macrocycle sampling algorithm that includes surrounding receptor atoms to restrict conformational search space
    • Updated macrocycle_sample.py script replaces macro_sample.py
  • Improved handling of ring nitrogen atom substituents during ring template conformation generation
  • tug_align.py now supports 2D ligand files as inputs
  • tug_align.py exposes several new command-line options that allow finer control of the alignment convergence criteria which are useful in reducing convergence times for large molecules such as cyclic peptides

Medical Chemistry Design

Ligand Designer

  • Enable user-specificed or automatically generated ligand protonation and tautomeric states from the 2D/3D editing workflow for MCS Docking

Biologics Drug Discovery

  • Completely rewritten MMGBSA Residue Scanning backend that is more reliable and has improved support for a wider variety of non-standard amino acids and mutation of DNA/RNA
  • New high-throughput, machine learning-based antibody and nanobody structure prediction with ImmuneBuilder. With throughput of about one minute per structure, it is suitable for batch modeling of thousands of structures
  • Automatically identify and annotate for visualization TCR alpha and beta chains from a FASTA file (commandline run_tcr_modeling.py)
  • Updated N-glycosylation PROSITE pattern that is less restrictive

Materials Science

GUI for Quantum ESPRESSO

Product: Quantum ESPRESSO (QE) Interface

  • Workflow solution to calculate the defect energy
  • Workflow action menu (WAM) for output from periodic DFT convergence test
  • Support for setting total magnetization/charge for each structure
  • Support for computing thermodynamic properties via dynmat.x (command line)
  • Support for stopping an NEB calculation and returning intermediate structures

Materials Informatics

Product: MS Informatics

  • Formulation ML: Option to control advanced settings
  • Machine Learning Property: Skip structures outside the model scope
  • Machine Learning Property: Prediction of singlet (S0) to triplet (T1) energy
  • Machine Learning Property: Prediction of hole / electron reorganization energy
  • Machine Learning Property: Prediction of orbital (HOMO / LUMO) energy
  • Machine Learning Property: Updates to existing models

Coarse-Grained (CG) Molecular Dynamics

Product: MS CG

  • Automated CG Mapping: Mapping MARTINI for speciality chemicals / polymers
  • CG FF Builder: Support for exporting atomistic reference data

Dielectric properties

Product: MS Dielectric

  • Complex Permittivity: Linear fitting parameter retained during exponential fitting

Reactivity

Product: MS Reactivity

  • Nanoreactor: Control over thermostat bath temperature
  • Nanoreactor: Support for parallelization of xTB dynamics simulations
  • Reaction Workflow: Support for the use of the xTB Hessian for transition states
  • Reaction Workflow: Use of input conformers when conformation search fails
  • Reaction Workflow: Improved SCF convergence for energy calculations

Microkinetics

Product: MS Microkinetics

  • Microkinetic Modeling: Improved speed/scalability by up to 2 orders of magnitude
  • Microkinetic Modeling: Improved model for multiple catalysts / catalyst site types
  • Microkinetic Modeling: Improved data visualization in the viewer
  • Microkinetic Modeling: Job name shown in the viewer

Reactive Interface Simulator

Product: MS RIS

  • Solid Electrolyte Interphase: Improved subjob queuing coordination
  • Solid Electrolyte Interphase: Support for constant pressure (NpT) simulations (command-line)

MS Maestro Builders and Tools

  • GUI panel to digitally design, manage, and visualize OLED devices
  • Single Complex: Improved UI for better usability
  • Solvate System: Support for generating multiple configurations by random seeds

Classical Mechanics

  • Workflow solution to compute thin plane shear friction
  • Barrier Potential for MD: Support for reading barrier information from entry
  • Evaporation: Option to plot total number of molecules removed from the system
  • Evaporation: Information tied to barriers (when applied) logged in the output
  • Polymer Crosslink: Improved speed by efficient checking for ring spears
  • Trajectory Density Analysis: Option to display multiple density depths
  • Trajectory Density Analysis: Export option for 2D heat-map plot

Quantum Mechanics

  • Workflow solution to plot phase diagrams based on energy
  • Workflow solution to compute and analyze computational ellipsometry data
  • QM Convergence Monitor: Easy access to the structure from the last step

Education Content

Life Science

  • New tutorial: Refining crystallographic protein-ligand structures using GlideXtal and Phenix/OPLS

Materials Science

  • New tutorial: Automated Martini Fitting for Coarse-Grained Simulations
  • New tutorial: Thin Shear
  • New tutorial: Defect Energy Calculation
  • New tutorial: Optoelectronics Device Designer
  • New tutorial: Computational Ellipsometry
  • New tutorial: Phase Diagrams
  • New tutorial: Ab initio Molecular Dynamics Simulations of Li-ion Diffusion in Solid State Electrolytes
  • Updated tutorial: Microkinetic Modeling
  • Updated tutorial: Organometallic Complexes

LiveDesign

What’s New in 2025-1

  • Biologics
    • Forms view now require the Sequence Viewer to drilldown from another widget
    • Performing a sequence alignment now shows an alert indicating the type of Alignment, and provides a “Try Again” button if the alignment fails
    • Forms views that contain both a Sequence Viewer and 3D Visualizer permit a synchronized selection of amino acid residues, such that selecting an amino acid in the Sequence Viewer will highlight the amino acid in the 3D Visualizer, and vice versa
    • Sequence search options now permit searching the Database, Active LiveReports, and Other LiveReports
    • View non-natural amino acid molecular structure in a tooltip by hovering over residues in the sequence viewer
    • Toggle monomers’ display format in the sequence viewer to view either single-letter FASTA format, or a custom symbol for non-natural monomers (up to 6 characters)
    • Reset Gap Penalties, Numbering Scheme, and Scoring Metrics to their default values for each alignment method
    • Substructures searches in Advanced search panel support a “Match by Child” option, which enables searching against an entity’s subcomponents
  • Performance Improvements
    • Small Molecules uploaded from file imports, enumeration, and Maestro uploads appear in the LiveReport more quickly
  • Landing Page: View all of a compound’s experimental data on the Compound’s detail page
  • Data and Columns Tree:
    • Group Multi-Parameter Optimization columns and Formulas columns into folders
    • The button text to create new Formulas, MPOs and FFCs has been changed from “NEW” to “CREATE”
    • Published Limited Assay Columns now show the [LIM] prefix in the Data & Columns Tree
  • A new MAE-FILE macro for Protocols in the Admin Panel allows Protocol and Models to access compounds and biologics using a Maestro file format. The Maestro file format includes residue information for Biologics
  • Configure LiveDesign to send an email on usage statistics, including: the number of unique logins over the last month, total number of active users, Number of compounds added last week, total number of compounds, total number of LiveReports updated last week, total number of active LiveReports and number of active LiveReports. For each user, the following information is reported: username, date of first login, date of last login, number of owned LiveReports
  • Configure a server-wide search setting to set the default searching behavior in the search panel to the Database, the Active LiveReport, or Other LiveReports
  • A new LDClient method, get_all_compounds(), enables retrieving all compounds within a specified list of projects
  • Sketched reactions within the Reaction Enumeration tool can now use atom queries to add greater specificity to the reaction definition

What’s Been Fixed

  • Updating a LiveReport template would create a duplicate template, and now correctly updates the template
  • Model columns that were used as inputs to more than 65,536 columns would fail to calculate values for new compounds, and now correctly calculate and display the predicted values
  • Pasting into the sketcher would fail on Windows when touchscreen was enabled, and now correctly pastes structures
  • LiveReports with hundreds of formula columns would show flashing cells, and slow down other LiveReports, but now correctly calculate and do not affect other LiveReports’ performance
  • Formulas that used the combine() function could not be used as input to subsequent formulas, and the formula would fail to calculate. Formulas using the combine() function now correctly calculate when they are input to other formulas
  • Selecting rows in the main spreadsheet would not show the 3D structure in the 3D Visualizer, and now correctly shows the 3D structure
  • LiveReports would show red error bars when multiple input values to a parameterized model changed simultaneously in the spreadsheet, and now the LiveReport loads correctly
  • The tautomer deduplication logic previously deduplicated keto-enol tautomers, but caused an unintended side effect of deduplicating compounds that were not tautomers. The updated tautomer deduplication logic has been reverted to the old behavior found in LiveDesign version 2024-4 and earlier
  • Popping out a model column’s cell that contained and image would open two tabs in the browser (one tab with the image, and one blank tab), and now only opens a tab with the image
  • Previously, changes to gap penalties were not persisting when saved in the Forms view, and now gap penalty values will persist correctly upon modification and saving
  • The “Batch Create Limited Assay Columns” option previously was unavailable in the column menu, if the column was within a column group, and now the option always appears
  • The residue synchronization between the 3D Visualizer and Sequence Viewer now remains functional even after changing the numbering scheme in Sequence Viewer
  • Model results would occasionally appear as Failed in the LiveReport, when in fact the model ran successfully, and now model results correctly show results in the LiveReport
  • The Assay Viewer Tool would occasionally change the date filter to 0 days and would not show data, and now defaults to showing data that was uploaded within the previous day.
  • Model results would occasionally appear as Failed in the LiveReport, when in fact the model ran successfully, and now model results correctly show results in the LiveReport
  • Ligand Designer’s deleted poses would occasionally reappear, and now no longer reappear
  • Protocols and Models showed their created date as one day earlier than their actual created date, and now show the correct date
  • LiveReport filters did not support filtering for Real or Virtual Biologics, and now provide quick toggles to apply those filters
  • Opening a model attachment from the main spreadsheet (e.g., a LID from a Glide model) would fail to show the image, and now correctly shows the image
  • Plot tooltips could not be dragged and moved after pinning to the screen, and now can be dragged to a new position after pinning
  • Importing a biologic entity would occasionally not show its subcomponents in the main spreadsheet, and now will correctly show its subcomponents
  • LiveDesign would occasionally fail to open LiveReports due to a database lock, and now no longer will freeze
  • Filtering out a frozen row would show flashing squares in the first row in the main spreadsheet, and now correctly shows that row’s data
  • The sequence viewer would show all entities in the LiveReport, even when only one entity was selected, and now correctly shows only the selected entities
  • Horizontally scrolling the sequence viewer would reset the ruler, and show the wrong residue numbers at the top, and now shows the correct residue numbers
  • Opening the Project Picker would occasionally take several seconds to show the list of projects, and now shows the list of projects immediately
  • LiveDesign would occasionally fail to open LiveReports due to a database lock, and now no longer will freeze
  • Filtering out a frozen row would show flashing squares in the first row in the main spreadsheet, and now correctly shows that row’s data
  • Exporting a LiveReport defaulted to exporting a subset of columns, and now defaults to exporting all of the columns
  • Syncing LiveDesign users with external authentication systems would fail because LiveDesign counted unlicensed users against the total license count, and now permits syncing users as long as there are available licenses
  • License files would occasionally fail to upload in the Admin Panel UI, and now correctly upload
  • User email addresses would not save in LiveDesign when using Single Sign-on, and LiveReport notifications would not be emailed to users. Email addresses and are now correctly saved
  • LiveReport tabs would disappear after logging out and logging in, and now correctly appear after logging back in
  • Duplicating a LiveReport defaulted to copying a subset of columns, and now defaults to copying all of the columns

Training & Resources

Online Certification Courses

Level up your skill set with hands-on, online molecular modeling courses. These self-paced courses cover a range of scientific topics and include access to Schrödinger software and support.

Tutorials

Learn how to deploy the technology and best practices of Schrödinger software for your project success. Find training resources, tutorials, quick start guides, videos, and more.

Other Resources

17th Winter Conference on Medicinal & Bioorganic Chemistry

Conference

17th Winter Conference on Medicinal & Bioorganic Chemistry

CalendarDate & Time
  • February 2nd-6th, 2025
LocationLocation
  • Steamboat Springs, Colorado

Schrödinger is excited to be participating in the 17th Winter Conference on Medicinal & Bioorganic Chemistry taking place on February 2nd – 6th in Steamboat Springs, Colorado. Join us for a poster presentation by Gang Wang, Principal Scientist II at Schrödinger, titled “Pairing machine learning and FEP+ enables the discovery of P38-MK2 molecular glues.”

icon time FEB 4 | 7:00PM
Pairing machine learning and FEP+ enables the discovery of P38-MK2 molecular glues

Speaker:
Gang Wang, Principal Scientist II, Schrödinger

Abstract:
Mitogen activating protein kinases (MAPK)-activated kinase MK2 phosphorylates AU-rich element binding proteins (AUBPs) that modulate the stability of cytokine mRNA. An emerging class of inhibitors create a ternary complex with upstream kinase p38 and MK2. These molecular glues demonstrate superior properties relative to orthosteric inhibitors of either p38 or MK2 but their mechanism of action presents challenges to computationally driven design. Herein we describe our computational design strategy, including the use of FEP+ coupled with Machine Learning (e.g., AutoDesigner and AL-FEP+), to discover multiple structurally diverse, potent, and highly selective molecular glues. Compounds 1 and 2 demonstrate pronounced reduction in TNFα levels after PO dosing in LPS mouse models and represent the validation of our modeling workflow for challenging molecular glues.

Schrödinger advances materials informatics for faster development of next-gen composites

Schrödinger advances materials informatics for faster development of next-gen composites

Overview

Cutting time to market by multiple orders of magnitude, machine learning and physics-based approaches are combined to open new possibilities for innovations in biomaterials, fire-resistant composites, space applications, hydrogen tanks and more.

Read full article

High-performance materials discovery: A decade of cloud-enabled breakthroughs

FEB 26, 2025

High-performance materials discovery: A decade of cloud-enabled breakthroughs

Schrödinger is excited to be hosting a webinar in collaboration with C&EN on February 26th. Join us for a presentation by Mathew D. Halls, Senior Vice President of Materials Science at Schrödinger, titled “High-performance materials discovery: A decade of cloud-enabled breakthroughs.”

Speaker:

Mathew D. Halls, Ph.D. Senior Vice President, Materials Science

Abstract:

Computational approaches combining physics-based molecular modeling with machine learning have revolutionized materials discovery at scale. But challenges still exist when it comes to large-scale and mega-scale simulations in terms of knowledge, infrastructure, speed, and resources.

This talk will showcase how Schrödinger’s integrated materials science platform enables massive parallel screening and de novo design campaigns across diverse applications. Through real-world case studies, we will demonstrate automated solutions that have successfully impacted R&D efforts across industries, including designing novel polymers, identifying promising hole-transport materials for organic electronics, and accelerating the discovery of organometallic precursors for thin film processing. We will describe how cloud computing infrastructure in tandem with Schrödinger’s 10+ years of experience executing and supporting large-scale projects facilitates unprecedented throughput in materials screening. Attendees will gain actionable strategies for implementing large-scale computational screening in their own materials research programs, along with best practices for integrating simulation with experiment.

Key Learning Objectives:

  • Learn to leverage advanced cloud computing infrastructure to meet the needs of large-scale materials simulations for industry applications
  • Hear case studies and customer success stories across industries including organic electronics, catalysis, energy capture and storage, polymeric materials, consumer packaged goods, pharmaceutical formulations, and thin film processing
  • Learn how Schrödinger can ensure a smooth implementation of a cloud-based materials modeling platform for your organization to maximize the value of digital simulations

Who Should Attend:

R&D Leaders
Innovation Managers
Digitization Managers
Synthetic Chemists
Materials Scientists
Chemical Engineers
Materials Research Engineers
Computational Chemists
Computational Materials Scientists

Our Speaker

Mathew D. Halls, Ph.D.

Senior Vice President, Materials Science, Schrödinger

Mathew D. Halls is responsible for leading the materials science program at Schrödinger. Mat received his Ph.D. in 2001 from Wayne State University in Detroit, MI, under the supervision of Professor H. Bernhard Schlegel. Over the past 20 years, Mat has worked with Fortune 500 companies to advance the adoption of atomic-scale materials simulation and machine learning in diverse industries including aerospace, electronics and specialty chemicals. He has made significant research contributions in areas such as computational spectroscopy, organic optoelectronic materials, nanocarbon-polymer interfaces, thin-film precursors and deposition processes and battery electrolyte additives; with his work being cited more than 7500 times.