All Schrödinger offices worldwide will be closed for the week of August 17-21 as part of a company-wide initiative to rest and recharge. Please expect limited responses during this time. Scientific and Technical Support team members will be available to answer emergency support issues only.

Accelerating Product Development: The Industrial Shift to AI/ML-Driven Formulation

Accelerating Product Development: The Industrial Shift to AI/ML-Driven Formulation

Schrödinger participated in a podcast hosted by Innovation Research Interchange on September 18th.

In this discussion, we explored the rapidly evolving role of modeling and machine learning in formulation design; from a supplementary tool to a driving force of innovation. Once considered a “nice to have,” computational modeling is now helping to replace costly and time-consuming physical experimentation, and accelerating product development across industries from CPG to aerospace to semiconductor.

We discussed how advances in AI/ML and physics-based simulations are enabling researchers to tackle the growing complexity of real-world formulations, bridging the gap between theory and commercial products. Whether you’re a formulation scientist, data enthusiast, or R&D leader, this discussion sheds light on how digital tools are transforming the lab and the future.

Our Speaker
Jeffrey Sanders

Product Manager and Scientific Lead, Consumer Goods, Schrödinger

Jeff Sanders received his B.S. in applied physics from Worcester Polytechnic Institute and then his Ph.D. in biophysics and molecular pharmacology from Thomas Jefferson Medical College. Since joining Schrödinger in 2013, he has served several roles. Jeff is currently the product manager and technical lead for the consumer packaged goods applications group. Additionally, he is a managing board member of the Food Engineering, Expansion, and Development (FEED) Institute, and also holds a faculty position in the Food Science Department at UMass Amherst.

Hit discovery course bundle

Hit Discovery Course Bundle_Hero

Hit discovery course bundle

Includes access to all paid, intermediate life science courses, including: designing quality ligand libraries, target enablement, validation, and preparation, and virtual screening with integrated physics and machine learning

Details
Available Languages
Chinese, English, Japanese, Korean
Duration
3 months from selected start date
Level
Intermediate
Cost
$1510 for non-student users
$525 for student / post-doc
Who should take this course?
Medicinal chemists, cheminformaticians, ML scientists, new computational chemists

Overview

As structural data and ligand libraries continue to grow, so does the demand for validated, scalable, and computationally efficient hit discovery workflows. 

This course bundle combines all three of Schrödinger’s intermediate life science courses into one powerful program, designed to build practical expertise across the entire virtual screening pipeline.

Through this series you’ll gain hands-on experience with Schrödinger’s industry-leading Maestro and command-line interface. Ideal for scientists looking to enhance their practical skills in structure-based modeling, ligand library design, and virtual screening techniques. 

An opportunity to professionally develop, deepen drug discovery skills, and earn certifications and digital badges that demonstrate capabilities across all major stages of the target validation and hit discovery process.

  • Prepare and refine protein structures, including exercises with AlphaFold structures and cryptic pocket identification
  • Understand the vastness of chemical space, and design and filter ligand libraries using profiling and enumeration strategies
  • Execute virtual screening campaigns with Active Learning Glide and other advanced computational tools
  • Work through real-world case studies challenging your skills in each focus area
  • Learn on your own schedule with expert guidance and curated learning content


This course comes with temporary access to a web-based version of Schrödinger software, complete with licenses and compute resources

Requirements
  • A computer with reliable high speed internet access (8 Mbps or better)
  • A mouse and/or external monitor (recommended but not required)
  • Working knowledge of general chemistry
  • Working knowledge of Maestro. Please work through our Getting Going with Maestro resources to become familiar with using Maestro.
  • (Optional) Prior completion of the Introduction to molecular modeling in drug discovery online certification course.
Certification
  • A certificate signed by the Schrödinger course lead to add to your CV or resume
  • A badge that can be posted to social media, such as LinkedIn
background pattern

What you will learn

Target enablement and assessment

Prepare, assess, and refine experimental (X-ray, cryo-EM) and ML-predicted (AlphaFold, homology) structures. Identify druggable binding sites and characterize cryptic pockets

Library design

Explore chemical space, profile vendor libraries, and generate tailored in silico enumerated libraries. Learn strategies to filter libraries to remove liabilities while preserving diversity

Executing and validating virtual screens

Run pilot virtual screens, prepare receptor grids, apply known active enrichment validation techniques

Hit evaluation and prioritization

Evaluate and prioritize hits, including inspection and clustering. Scale screening with machine learning and AB-FEP+

Course syllabus

The course bundle includes access to the following three courses in their entirety during the single course session.

Designing quality ligand libraries
Target enablement, preparation, & validation
Virtual screening with integrated physics & machine learning

Need help obtaining funding for a Schrödinger Online Course?

We proudly support the next generation of scientists and are committed to providing opportunities to those with limited resources. Learn about your funding options for our online certification courses as a student, post-doc, or industry scientist and enroll today!

What our alumni say

“It is wonderful to have a group of high-quality professors teaching you computational chemistry. I think that the creation of ligand libraries in silico is an extremely useful skill in many circumstances. As a computational biologist it is gratifying to have this skill in my curriculum. Very grateful for the whole process, as always Schrödinger the best.”
Andres. R. Ch. PradaBiologist. M.Sc. Biostatistics. M.Sc. Computational Biology. Master in Molecular Biology, Andes University
“As a Data Engineer with a general background in software engineering, this course has had a hugely positive impact on my ability to collaborate with computational chemists. This course provided a valuable balance of theory and practice, giving me the scientific context necessary for making technical decisions when designing pipelines for my colleagues.”
Lillian Campbell
Lillian CampbellData Engineer, Oddity Labs
“The course was designed to tackle the pressing need of drug discovery acceleration when high precision protein prediction methods are readily available. I highly recommend this course.”
Wei WangAssistant Professor, Icahn School of Medicine
“The course was easy and enjoyable to follow, with background information that gives insight into how each technique is valued in modern drug discovery research. I now feel like an adept Maestro user and can steer my career towards molecular modeling with confidence.”
Joseph EganIntern, center of medicine research and innovation
“I had an amazing experience with the virtual working station! As a PhD candidate with a background in medicinal chemistry, the knowledge and skills I acquired in this course will benefit me in reaching my career goal.”
Sumaiya NahidGraduate Research Assistant, University of Nebraska Medical Center (UNMC)
“The course provided a comprehensive understanding of virtual screening methodologies that helped streamline my drug discovery process. It had a direct impact on the progress and success of my research projects.”
Abdulbasit Haliru YakubuPhD student, University of Southampton

Show off your newly acquired skills with a course badge and certificate

When you complete a course with us in molecular modeling and are ready to share what you learned with your colleagues and employers, you can share your certificate and badge on your LinkedIn profile.

Frequently asked questions

How much does the Hit discovery course bundle online course cost?

Pricing varies by each course and by the participant type. For students wishing to take this, we offer a student price of $500, and $1435 for non-students.

What time are the lectures?

Once the course session begins, all lectures are asynchronous and you can view the self-paced videos, tutorials, and assignments at your convenience. When registering for the course you will select the start and end date. Within those dates, you will have asynchronous access to the course material and virtual workstation to work on the course when it best suits your schedule.

How could I pay for this course?

Interested participants can pay for the course by completing their registration and using the credit card portal for an instant sign up. Please note that a credit card is required as we do not accept debit cards. Additionally, we can provide a purchase order upon request, please email online-learning@schrodinger.com if you are interested in this option. If you have any questions regarding how to pay for the course, please visit our funding options page.

Are there any scholarship opportunities available for students?

Schrödinger is committed to supporting students with limited resources. Schrödinger’s mission is to improve human health and quality of life by transforming the way therapeutics and materials are discovered. Schrödinger proudly supports the next generation of scientists. We have created a scholarship program that is open to full-time students or post-docs to students who can demonstrate financial need, and have a statement of support from the academic advisor. Please complete the application form if you qualify for our scholarship program!

Will material still be available after a course ends?

While access to the software will end when the course closes, some of the material within the course (slides, papers, and tutorials) are available for download so that you can refer back to it after the course. Other materials, such as videos, quizzes, and access to the software, will only be available for the duration of the course.

Do I need access to the software to be able to do the course? Do I have to purchase the software separately?

For the duration of the course, you will have access to a web-based version of Maestro, Bioluminate, Materials Science Maestro and/or LiveDesign (depending on the course). You do not have to separately purchase access to any software. While access to the software will end when the course closes, some of the material within the course (slides, papers, and tutorials) are available for download so that you can refer back to it after the course. Other materials, such as videos, quizzes, and access to the software, will only be available for the duration of the course. Please note that Schrödinger software is only to be used for course-related purposes.

Related courses

Target Enablement Course Image_Edu_R1-2Website_Featured Image Life Science Life Science
Target enablement, preparation, & validation

Enabling protein structures from x-ray crystallography, cryo-EM, ML-methods, and homology modeling for structure-based computational workflows

Designing Quality Ligand Libraries_Featured image Life Science Life Science
Designing quality ligand libraries

Exploring chemical space, profiling and tailoring ligand libraries, validating docking models, and methods of enumeration for hit discovery

Virtual screening Life Science Life Science
Virtual screening with integrated physics & machine learning

Acquire essential skills in next-generation virtual screening, integrating physics and machine learning for smarter hit identification

Computational drug design and chemo-informatics: a hands-on course at the University of Antwerp

JUN 15, 2023

Computational drug design and chemo-informatics: a hands-on course at the University of Antwerp

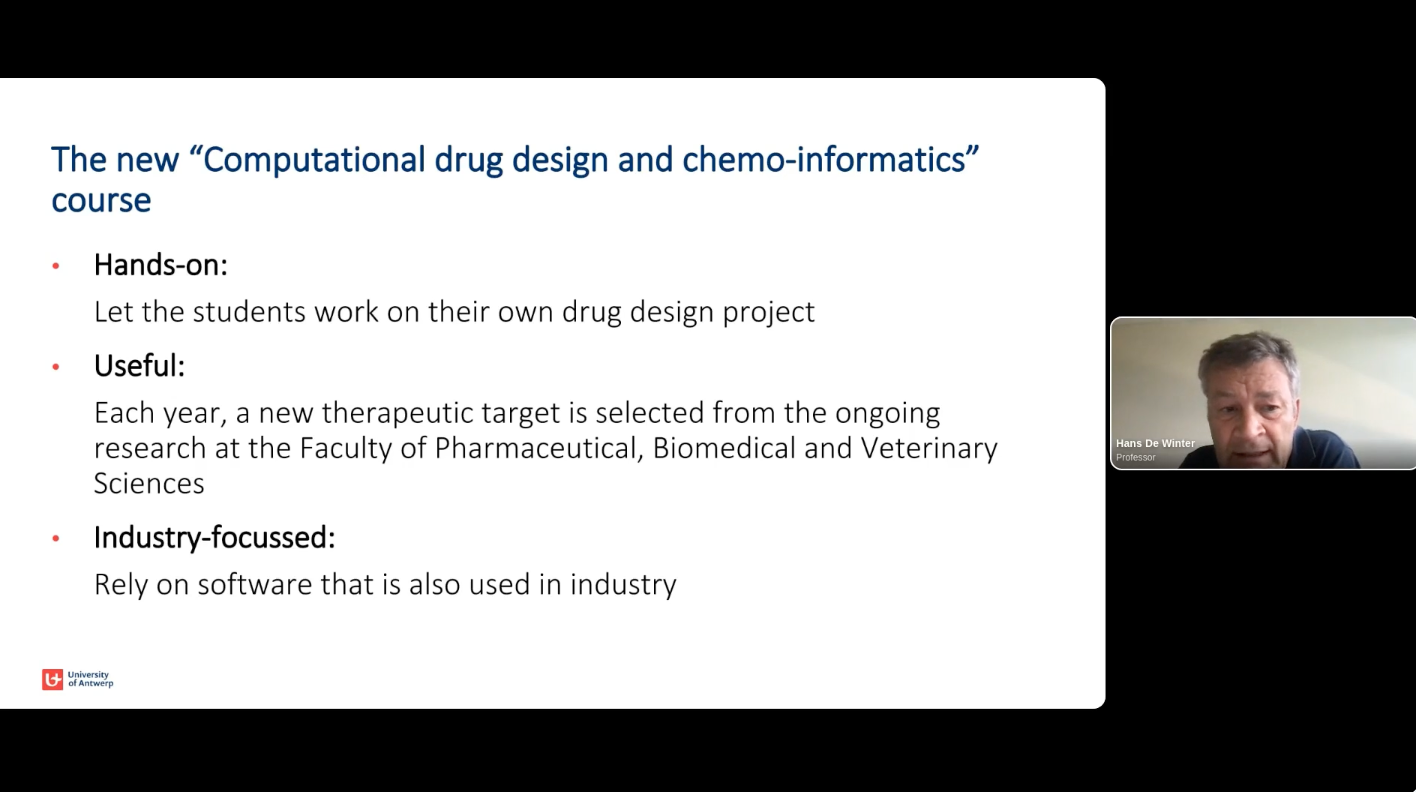
The University of Antwerp is the third-largest university in the Dutch-speaking region of Belgium, with over 20,000 students annually. Within the Biochemistry and Biotechnology curriculum, students have the option to take a three-ECTS course on computational drug design and chemo-informatics. The course is organized in a modular fashion and covers both theoretical and practical sessions.

During the theoretical sessions, students learn about chemo-informatics and virtual screening, which includes concepts such as chemical fingerprints, molecular similarity, clustering, machine learning models, and virtual screening performance metrics. The course also covers molecular docking and pharmacophore searching. The concepts covered in the theoretical sessions are then put into practice in a series of hands-on sessions.

For the chemo-informatics tasks, the students use Google Colab with RDKit as a chemo-informatics toolkit, while for the pharmacophore and docking-related aspects, they use Maestro, Phase, and Glide. These tools are made available through the “”Teaching with Schrödinger”” web-based virtual workstations, which allows students to access them from anywhere at any time. Finally, using an internally-developed virtual reality system, the students can graphically study the non-bonded interactions between ligand and protein.

At the start of the course, a drug design project is defined based on ongoing research programs in the Faculty. The goal of the project is to identify a limited number of commercially-available compounds (5-10) that are subsequently purchased and biochemically characterized for their inhibitory properties. The students complete the program with a written report, which serves as the basis for the oral examination at the end.

Our Speaker

Hans De Winter

University of Antwerp

Hans De Winter was appointed in 2013 as a professor of Computational Drug Design at the University of Antwerp (Belgium) after a long career in industry, first as a senior scientist at Johnson & Johnson in Beerse, Belgium, and subsequently as a co-founder and CSO of Silicos NV. He holds a PhD from the University of Leuven (Belgium) and completed post-doctoral stays at the Victorian College of Pharmacy (Australia) and the Rega Institute in Leuven (Belgium) before starting his career as a scientist in the pharmaceutical industry. Despite his elaborated industrial background during a period of more than 20 years, he has over 60 scientific publications and is listed as inventor on eight granted patents. Hans’ research interests are mainly situated in the field of computational medicinal chemistry and cheminformatics. Current research activities include: 1) molecular dynamics-based modeling of protein-protein interactions and unraveling the kinetics of lipid membrane-bound protein complexes using large-scale molecular dynamics calculations with Markov chain modeling; 2) the elaboration of spectrophore-based algorithms for exploration of ligand/protein interaction space; 3) the development of automated in silico ligand design systems. Additional focus points are the development of open-source and cloud-based interrogation software for large chemical and biological data repositories. Finally, his expertise in in silico drug design is used in several ongoing targeted drug discovery projects. Prof. Hans De Winter is the coordinating promoter or co-promoter of a number of PhDs and has a broad research network both in industry and in academia based on his expertise in modeling/chemoinformatics. He is also chairman of the Flemish computational chemistry division of the European Association of Chemical and Molecular Sciences (EuCheMS) since 2015. He teaches ‘organic chemistry’ to 1st year bachelor students in Pharmaceutical Sciences, and ‘computational drug design and cheminformatics’ to 1st year master students in Biochemistry at the University of Antwerp.

MS Formulation ML

MS Formulation ML

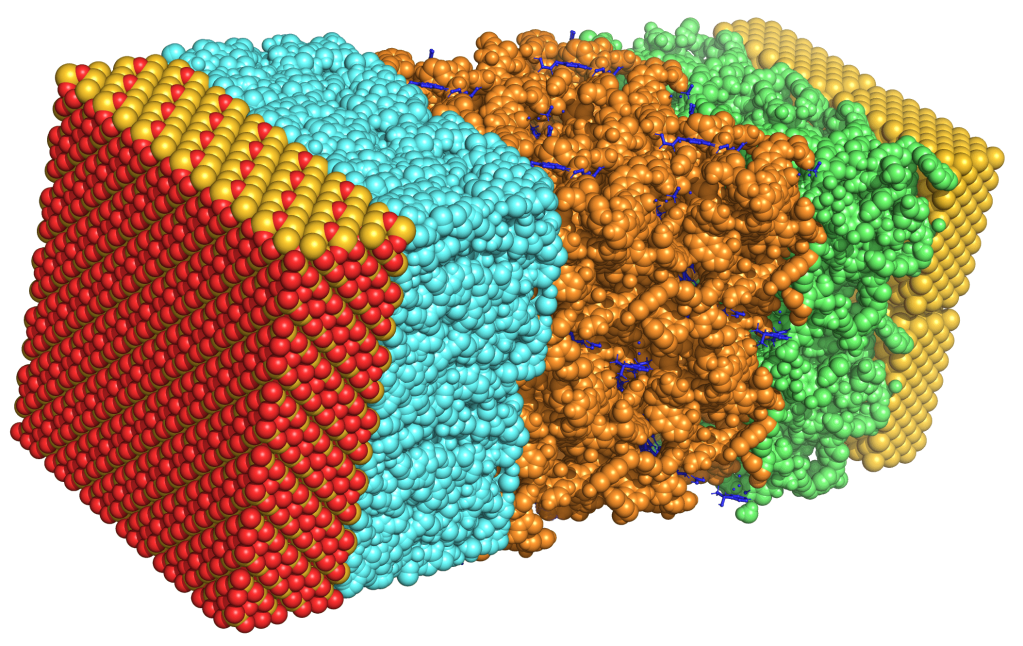
Automated machine learning solution to generate accurate formulation-property relationships and screen new formulations with desired properties

MS Formulation ML

Create accurate machine learning models to design better formulations

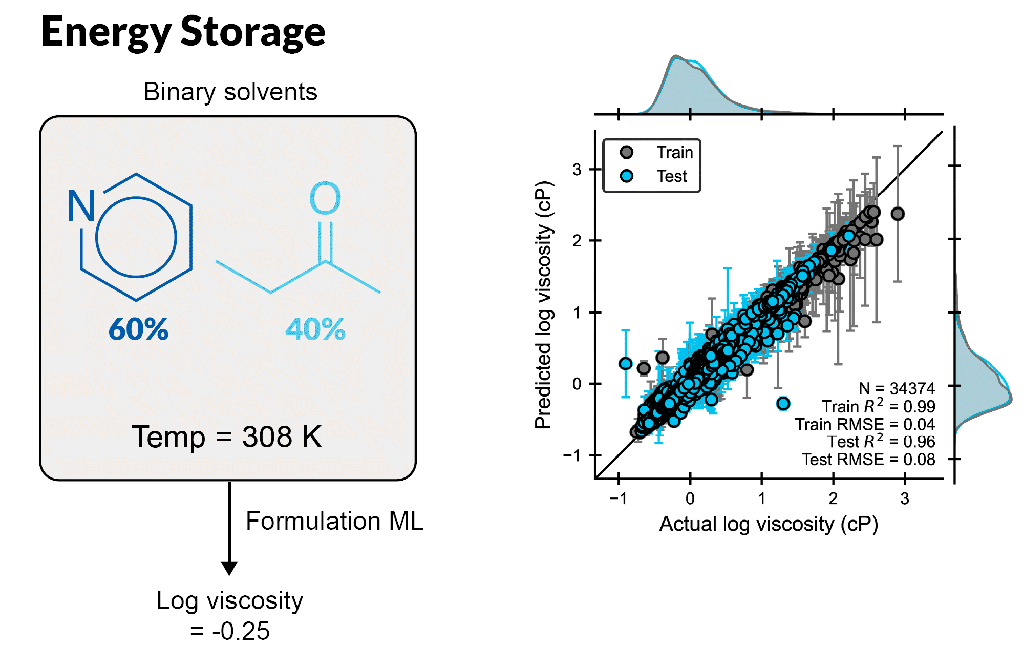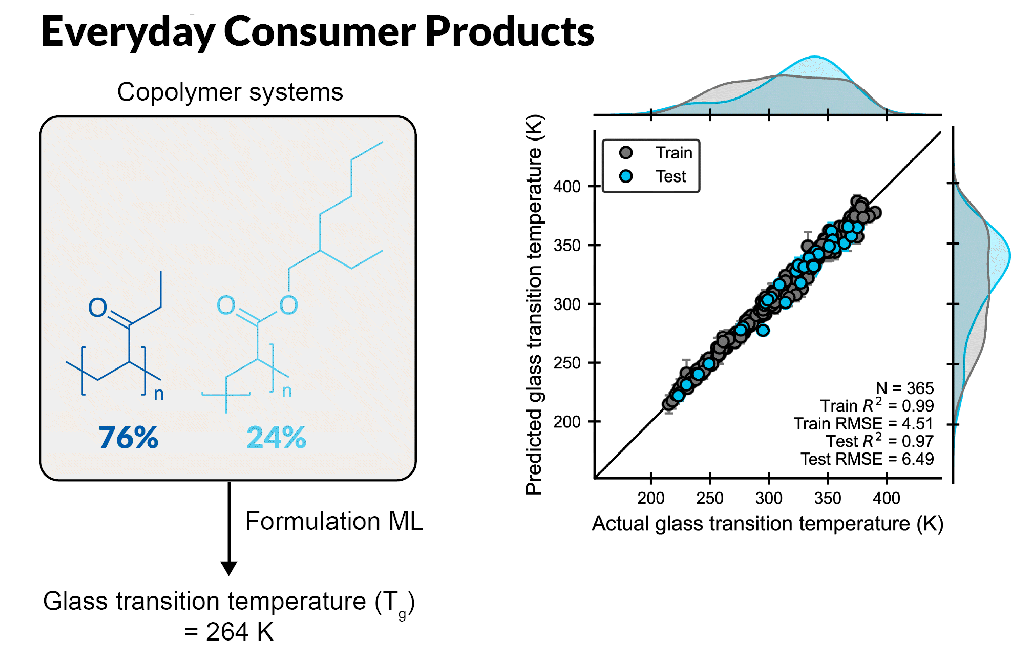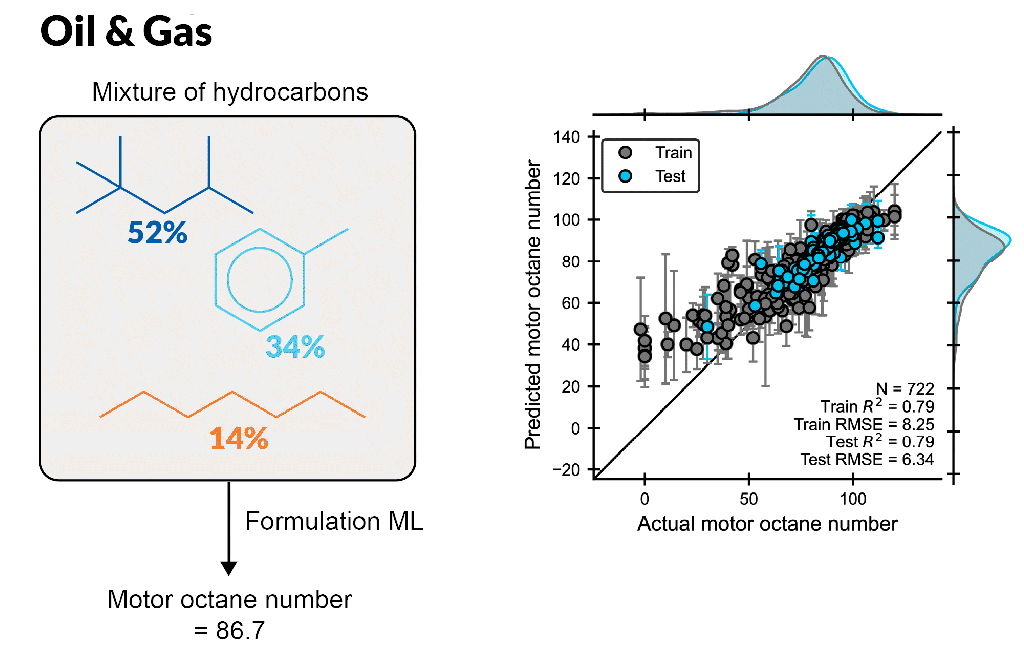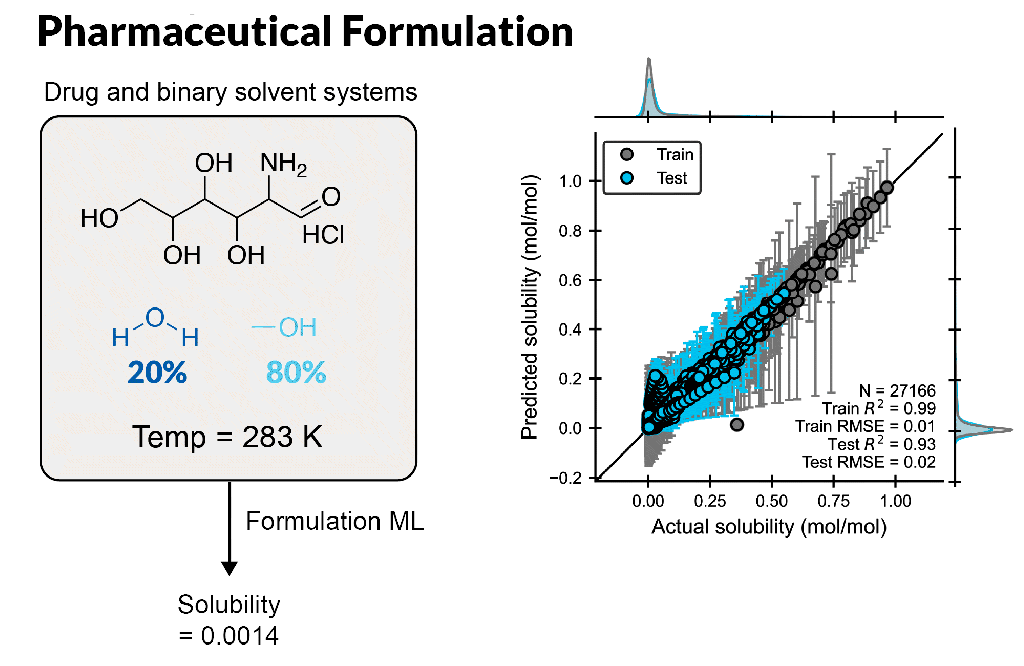
Formulation ML allows scientists to predict properties based on ingredient structures and compositions. Whether you are a formulation expert or just learning in this area, this automated, supervised learning solution enables you to gain deeper insight into formulation-property relationships.

Key Capabilities

Build formulation-property models for chemical mixtures with varying ingredient structures and compositions, which are scalable up to 100 ingredients or more
Rapidly predict novel formulations with new chemistry and composition, requiring only seconds per formulation
Understand which molecular features to focus on to fine-tune properties, leveraging feature importance tools to identify key descriptors for a property using a trained model
Enable accurate ML model development using expert cheminformatic descriptors and automatic hyperparameter tuning with minimal ML expertise
Input customized descriptors, including experimental data, in CSV format into the ML model to improve model performance
Optimize multiple properties simultaneously by modulating ingredient structure and compositions with trained ML models, providing suggestions of best formulations for the next experiment

Featured Resources

Materials Science Informatics Webinar Materials Science
AI/ML meets physics-based simulations: A new era in complex materials design

In this webinar, we demonstrate the application of this combined approach in designing materials and formulations across diverse materials science applications, from battery electrolytes and fuel mixtures to thermoplastics and OLED devices. 

Accelerating pharmaceutical formulations using machine learning approaches Webinar Life Science Materials Science
Accelerating pharmaceutical formulations using machine learning approaches

In this webinar, we will demonstrate how Schrödinger’s integrated ML- and physics-based approaches are transforming pharmaceutical formulation design.

Complex Formulations

Tutorial

Machine Learning for Formulations

Related Products

Desmond

High-performance molecular dynamics (MD) engine providing high scalability, throughput, and scientific accuracy

Jaguar

Quantum mechanics solution for rapid and accurate prediction of molecular structures and properties

DeepAutoQSAR

Automated, scalable solution for the training and application of predictive machine learning models

MS Informatics

Automated machine learning tools for materials science applications

MS Force Field Applications

Cutting-edge force field technologies for accurate property predictions

Publications

Leveraging high-throughput molecular simulations and machine learning for the design of chemical mixtures, Alex, C., et al. npj Comput Mater 11, 72, 2025, https://doi.org/10.1038/s41524-025-01552-2.

Schedule a consultation on Schrödinger’s Formulation ML

Contact us today to explore how you can leverage advanced simulation and AI/ML to transform formulation decisions and gain competitive advantage in your industry.

Don’t see your areas of interest in the current lists above? Reach out so we can help.

Form submitted

Thank you, we’ll be in touch soon.

Software & services to meet your organizational needs

Software Platform

Deploy digital materials discovery workflows with a comprehensive and user-friendly platform grounded in physics-based molecular modeling, machine learning, and team collaboration.

Research Services

Leverage Schrödinger’s expert computational scientists to assist at key stages in your materials discovery and development process.

Support & Training

Access expert support, educational materials, and training resources designed for both novice and experienced users.

OLED Device ML

OLED Device ML

Machine learning solution to investigate relationships between the architecture and performance of OLED devices for accelerated screening

OLED Device ML

Create machine learning models to enable high-throughput design and optimization of OLED devices

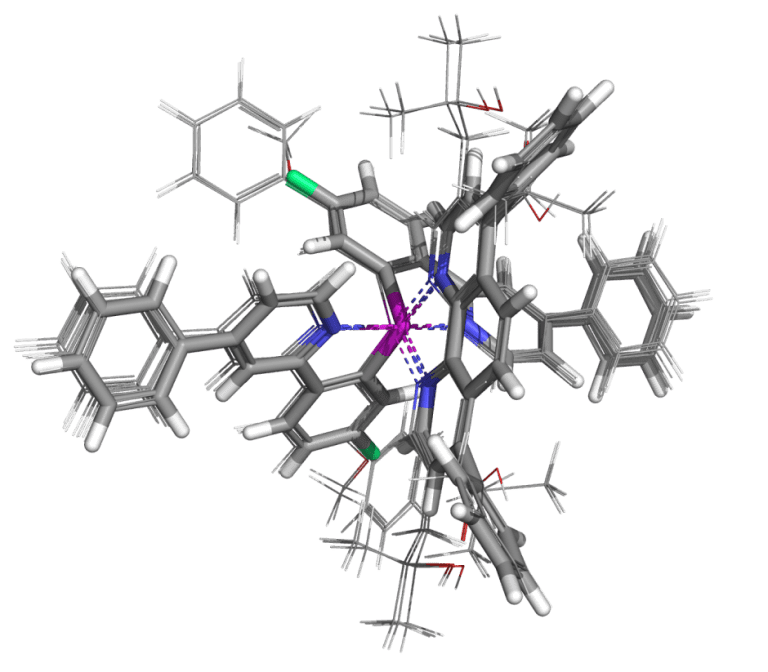
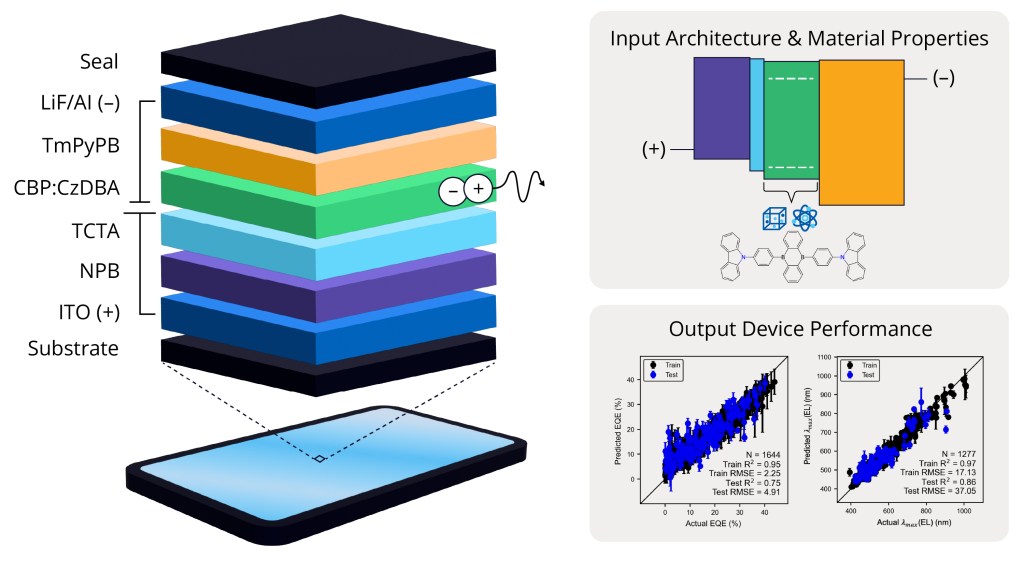
The OLED Device ML solution enables scientists to predict performance metrics that quantify the operational output, efficiency, and stability of multicomponent layered organic light-emitting diodes (OLEDs). These predictions are based upon simple and direct descriptions of device operation and architecture, such as the arrangement and chemical composition of layers. This offers a scalable solution for OLED developers seeking to perform targeted evaluations of device capability across novel design spaces.

Key Capabilities

Train chemistry-informed ML models to predict performance properties for OLED devices with varying layer arrangements and chemical compositions
Rapidly predict the performance of novel device structures to establish interpretable relationships between functionality and layer architectures and chemistry
Use pre-trained ML models to predict six different device performance metrics including external quantum efficiency, current efficiency, power efficiency, electroluminescence maximum peak position, electroluminescence bandwidth, and color of the emitted light
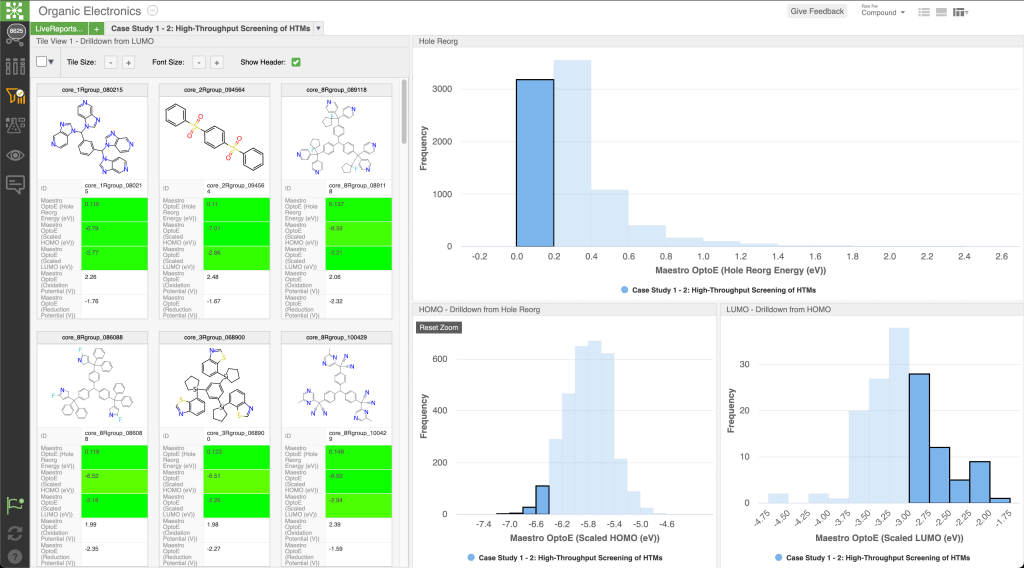
Benefit from an intuitive graphical interface that allows easy design and exploration of novel chemistry and device architectures, facilitated by the visualization of energy level diagrams with out-of-the-box QM descriptors and ML models

White paper

LiveDesign for Organic Electronics

Broad applications across materials science research areas

Related Products

MS Informatics

Automated machine learning tools for materials science applications

DeepAutoQSAR

Automated, scalable solution for the training and application of predictive machine learning models

Schedule a consultation on Schrödinger’s OLED Device ML

Contact us today to explore how you can leverage advanced simulation and AI/ML to design better electronic devices.

Don’t see your areas of interest in the current lists above? Reach out so we can help.

Form submitted

Thank you, we’ll be in touch soon.

Software & services to meet your organizational needs

Software Platform

Deploy digital materials discovery workflows with a comprehensive and user-friendly platform grounded in physics-based molecular modeling, machine learning, and team collaboration.

Research Services

Leverage Schrödinger’s expert computational scientists to assist at key stages in your materials discovery and development process.

Support & Training

Access expert support, educational materials, and training resources designed for both novice and experienced users.

Schrödinger デジタル創薬セミナー: Into the Clinic ~計算化学がもたらす創薬プロセスの変貌~ 第14回

NOV 19, 2024

Schrödinger デジタル創薬セミナー 14:
Advancements in machine learning enhanced in silico design: Impact on a pipeline of drug discovery programs

分子特性のシミュレーションは、物理ベースのアプローチを使用することで、構造と特性の関係に関する洞察を提供し、新薬の設計を支援する分野で長らく成功を収めてきました。近年では、AIや機械学習(ML)が物理ベースのモデリング技術と組み合わさり、革新の加速に大いに貢献しています。物理ベースのモデリングの精度と一般化能力が、AI/MLモデルのパフォーマンスを向上させ、データが少ない領域でも効果的に使用できるようにしています。逆に、AI/MLのスピードと柔軟性は、物理ベースのモデルが抱える時間的・空間的な限界を克服する手助けをし、予測精度と計算効率の両方を最適化する相乗効果を生み出します。

このウェビナーでは、機械学習を活用して創薬プログラムを推進する、以下の応用例について議論します。

  • FEP+を使用したアクティブラーニングによる、大規模なインシリコフラグメントスクリーニングでのヒット探索
  • インテリジェントな分子コア設計のためのde novoデザインワークフローの適用
  • インタラクティブなMLダッシュボードを用いたリード最適化におけるADMETプロファイルの強化のための実験データの活用

Our Speaker

Karl Leswing

Vice President Machine Learning, Schrödinger

Karl Leswing is the Vice President for Machine Learning at Schrödinger. In this role he oversees the research and execution of machine learning applications for Schrödinger’s digital chemistry platform. In 2017 he was a visiting researcher at the Pande Lab working on using deep learning techniques for drug discovery. During that time he co-authored MoleculeNet, a benchmarking paper analyzing machine learning techniques for chemoinformatics. Karl received his undergraduate degree from the University of Virginia, and a Master’s in machine learning from Georgia Tech.