All Schrödinger offices worldwide will be closed for the week of August 17-21 as part of a company-wide initiative to rest and recharge. Please expect limited responses during this time. Scientific and Technical Support team members will be available to answer emergency support issues only.

Moving Beyond Spreadsheets: Rational Design of Materials Using Advanced Informatics and Machine Learning

AUG 17, 2021

Moving Beyond Spreadsheets: Rational Design of Materials Using Advanced Informatics and Machine Learning

Speaker

Yuling An
Product Manager

Summary

In this webinar, Schrödinger’s Dr. Yuling An will demonstrate that machine learning, which often ignores the underlying physics, and physics-based modeling, which may require intensive computing resources, can naturally complement each other to create not only predictive models but also new materials with desired properties over an extensive design space. The growing urgency to digitize and make use of existing data, both from experiments and from simulations, through machine learning, also heightens the need of materials informatics platforms, which bring together automated computational workflows with data analysis and collaboration to make materials innovation more efficient and successful. Examples include recent progress in de novo OLED materials design and prediction of molecular volatility using Schrӧdinger’s materials informatics platform, LiveDesign.
Active Learning Applications

background pattern

Active Learning Applications

Accelerate discovery with machine learning

Amplify discovery across vast chemical space

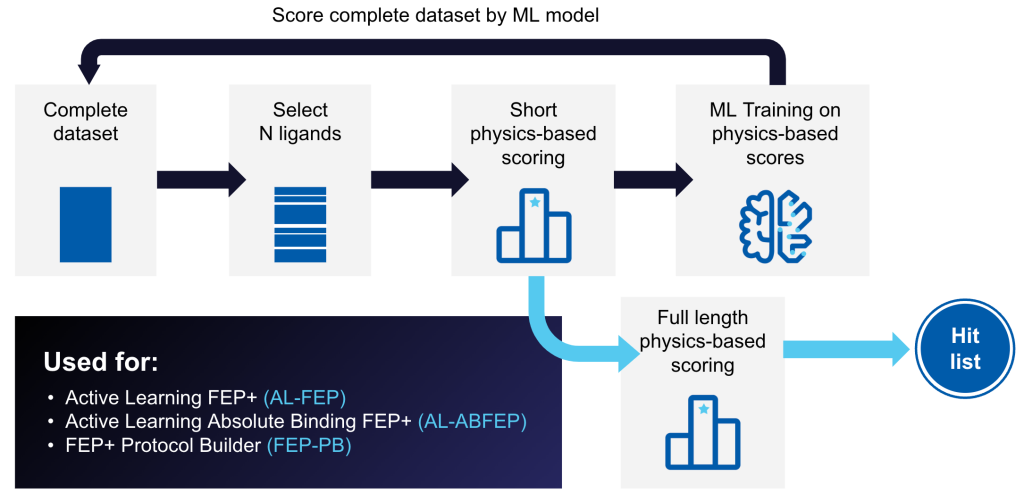
Active Learning Applications is a powerful tool that trains a machine learning (ML) model on physics-based data, such as FEP+ predicted affinities or Glide docking scores, iteratively sampled from a full library.

Trained models can rapidly generate predictions for new molecules and identify the highest-scoring compounds in ultra-large libraries at a fraction of the cost and speed of brute force methods.

Key applications across drug discovery

Active Learning Glide
Find potent hits in ultra-large libraries

Screen billions of compounds with Glide docking amplified by cutting-edge machine learning models in a fraction of the time. Use Active Learning to recover ~70% of the same top-scoring hits that would have been found from exhaustive docking of ultra-large libraries with Glide, for only 0.1% of the cost.

Active Learning FEP+
Explore diverse chemical space in lead optimization

Explore tens of thousands to hundred of thousands of idea compounds with Active Learning FEP+, against multiple hypotheses simultaneously, to quickly identify compounds that maintain or improve potency while achieving other design objectives.

FEP+ Protocol Builder
Expedite FEP+ use for challenging systems with a fully automated workflow

Rapidly generate accurate FEP+ protocols for systems that do not perform well with default settings. FEP+ Protocol Builder uses an Active Learning workflow to iteratively search the protocol parameter space to develop accurate FEP+ protocols, saving researcher time and increases the chances of successfully enabling FEP+.

Learn more

Active Learning Calculator

Glide (Dock All Compounds)

days
compute
cost

Active Learning Glide

Faster
days
compute
cost

For Compounds

Enter in the numbers for your project (type in the box or use slider) to compare compute time and cost.

*Estimated customer compute costs only, based on $0.06 per CPU hour and $.35 per GPU hour. Recommended hardware for AL-Learning Glide.
License costs are not included. Contact us for a quote.
We assume 1M of the best ligands are docked with the final model.

De Novo Design Workflow

Schrödinger’s De Novo Design Workflow is a fully-integrated, cloud-based design system for ultra-large scale chemical space exploration and refinement.

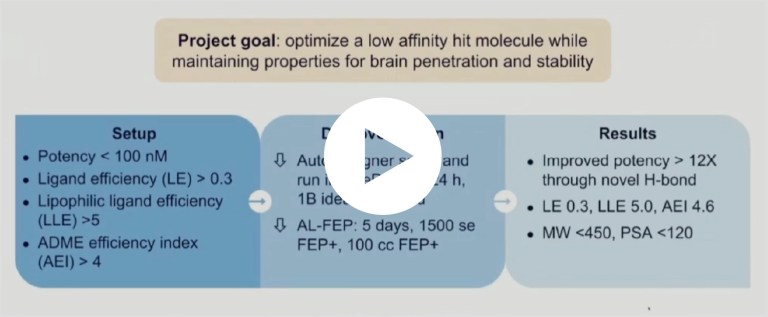
Starting from a hit molecule or lead series, the technology identifies synthetically tractable molecules that meet key project criteria by combining multiple compound enumeration strategies with an advanced filtering cascade (AutoDesigner) and rigorous potency scoring with free energy calculations (Active Learning FEP+).

Documentation & Tutorials

Get answers to common questions and learn best practices for using Schrödinger’s software.

Life Science Tutorial

Screening ultra-large libraries with Generative Glide

Screen billion-sized libraries in 24 hours with a generative ML approach.

Life Science Documentation

Active Learning Applications

Active Learning Glide documentation including online help and user manual.

Life Science Tutorial

Evaluating Large Ligand Libraries with Active Learning Glide

Set up a virtual screen to analyze a 1M ligand library from using Active Learning Glide.

Related Products

Learn more about the related computational technologies available to progress your research projects.

Glide

Industry-leading ligand-receptor docking solution

FEP+

High-performance free energy calculations for drug discovery

De Novo Design Workflow

Fully-integrated, cloud-based design system for ultra-large scale chemical space exploration and refinement

Publications

Browse the list of peer-reviewed publications using Schrödinger technology in related application areas.

Life Science Publication

Harnessing free energy calculations to achieve kinome-wide selectivity in drug discovery campaigns: Wee1 case study

Life Science Publication

Optimizing drug design by merging generative AI with a physics-based active learning framework

Life Science Publication

Active Learning FEP: Impact on Performance of AL Protocol and Chemical Diversity

Life Science Publication

FEP augmentation as a means to solve data paucity problems for machine learning in chemical biology

Life Science Publication

Pathfinder-Driven Chemical Space Exploration and Multiparameter Optimization in Tandem with Glide/IFD and QSAR-Based Active Learning Approach to Prioritize Design Ideas for FEP+ Calculations of SARS-CoV-2 PLpro Inhibitors

Life Science Publication

Discovery of a Novel Class of d-Amino Acid Oxidase Inhibitors Using the Schr’dinger Computational Platform

Life Science Publication

AutoDesigner, a De Novo Design Algorithm for Rapidly Exploring Large Chemical Space for Lead Optimization: Application to the Design and Synthesis of D-Amino Acid Oxidase Inhibitors

Materials Science Publication

Design of organic electronic materials with a goal-directed generative model powered by deep neural networks and high-throughput molecular simulations

Life Science Publication

Impacting Drug Discovery Projects with Large-Scale Enumerations, Machine Learning Strategies, and Free-Energy Predictions

Life Science Publication

Efficient Exploration of Chemical Space with Docking and Deep-Learning

Training & Resources

Online certification courses

Level up your skill set with hands-on, online molecular modeling courses. These self-paced courses cover a range of scientific topics and include access to Schrödinger software and support.

Tutorials

Learn how to deploy the technology and best practices of Schrödinger software for your project success. Find training resources, tutorials, quick start guides, videos, and more.

Accelerating the design and optimization of OLED materials using active learning

Accelerating the design and optimization of OLED materials using active learning

Introduction

OLEDs (Organic Light-Emitting Diodes) are extensively used in digital displays such as those in smart phones, television screens, computer monitors, and game consoles. They contain organic molecules with unique electronic structures that create light in the visible part of the spectrum through a process called electroluminescence. For an OLED to be commercially valuable, it should satisfy several constraints, including high efficiency, low-cost fabrication, and good electrochemical stability for long-term operation, making discovery of novel OLED materials a challenging problem.


Challenges

Molecular modeling and simulation tools have proven effective in materials discovery and are increasingly deployed in industrial R&D. Although digital simulations have offered tremendous time savings for R&D workflows compared to traditional experimental approaches, several challenges persist:

  • The potential chemical space for materials design and discovery is massive, even for highly constrained problems
  • Predictions based on density functional theory (DFT) calculations can be laborious and computationally expensive, limiting the number of candidates evaluated within fixed timescales and resources
  • It is challenging to maintain a high level of accuracy to properly assess the complexity of materials


Solution: Active Learning Workflows

A new approach is required to guide scientists to the best-performing or useful candidates. Thanks to the application of a machine learning (ML) paradigm called “active learning” (AL), Schrödinger has made this problem readily tractable. Recently, Schrödinger has developed active learning workflows which leverage the synergy between physics-based simulations and machine learning for optoelectronic properties predictions. The active learning workflow enables scientists to zoom in on the “best-performing” portion of a given sample space in a more efficient and cost-effective manner. The workflow allows:

  • Automated active learning calculations with minimum users input
  • Accounting for multiple optoelectronic parameters simultaneously for materials discovery
  • Combining active learning with DFT to efficiently identify materials with optimal properties
  • Employing built-in descriptors and fingerprints to featurize chemical structures
  • Building high-performance ML models using adaptive design procedures
  • Minimizing the number of time-consuming physics-based calculations
““The AL workflow enables accelerated design and discovery of optoelectronic materials. The workflow is fully automated, significantly reducing the number of physics-based simulations to predict materials properties in an extensive library. The rapid screening of datasets allows a better understanding of structure-function relationships for systematic design and application of optoelectronic materials with higher efficiency.””
Hadi Abroshan Senior Scientist, Schrödinger

Case Study

Recent studies by Schrödinger, published in Frontiers in Chemistry and presented at SID-Display Week 2022, have demonstrated the active learning paradigm for OLED materials discovery.1,2

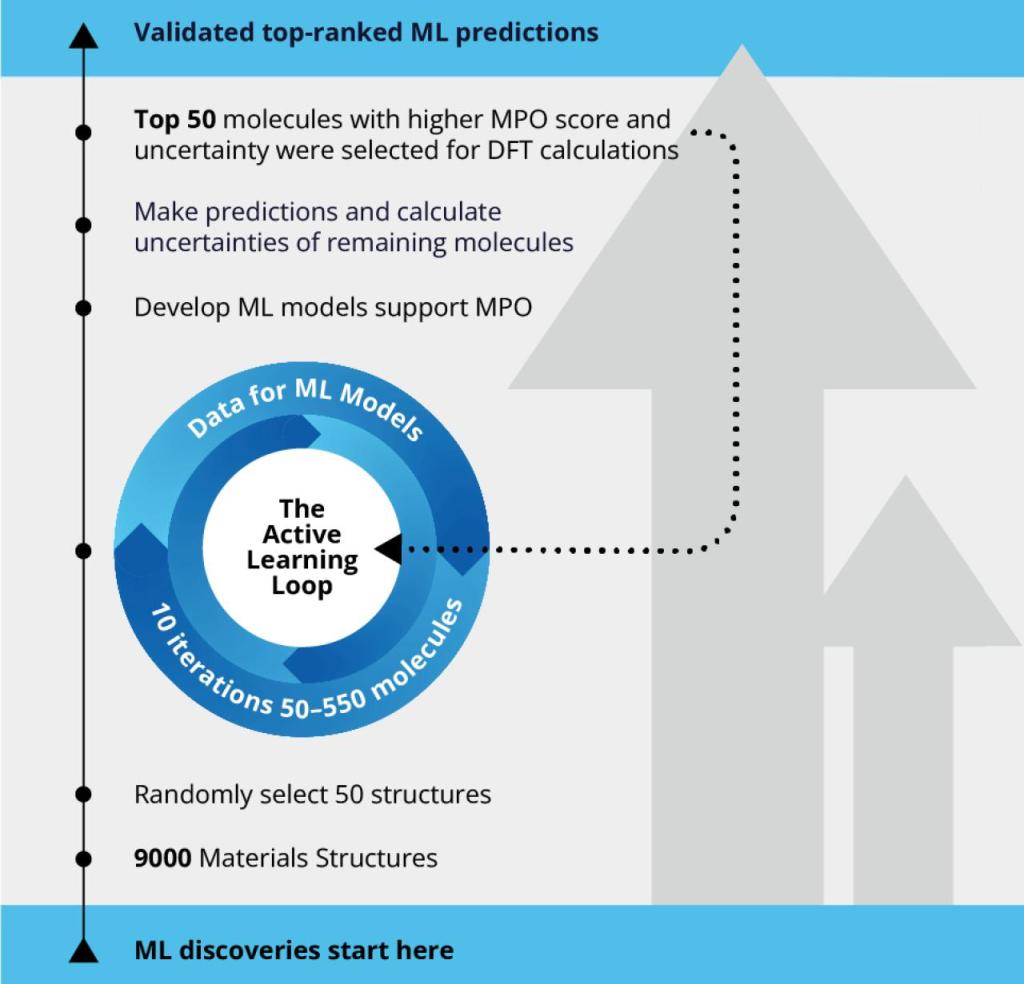
To explore the best-performing hole-transporting molecules for OLED, scientists screened a large pool of 9,000 molecules using the automated active learning workflow. The initial training set included 50 molecules for which a machine learning model was developed supporting efficient multi-parameter optimization (MPO). The ML model was then used to predict the rest of 8,950 molecules in the pool, each of which only costs a fraction of second. Top 50 molecules with higher MPO score and uncertainty were selected for DFT calculations and the calculated data were input for the next iteration of ML model training. The size of the training set for active learning was increased from 50 to 550 molecules in 10 iterations.

Active learning is the iteration of these steps until the DFT calculated data and the machine learning predicted data converge with sufficient accuracy. By using the AL approach, the scientific team was able to screen the chemical space of the materials library 18 times faster than the traditional approach of expensive quantum mechanical calculations, leading to considerable time and resource savings.


The Active Learning Workflow

Diagram of the automated active learning workflow for the design and discovery of novel OLED materials

References

  1. Active Learning Accelerates Design and Optimization of Hole Transporting Materials for Organic Electronics Hadi Abroshan, H. Shaun Kwak, Yuling An, Christopher Brown, Anand Chandrasekaran, Paul Winget and Mathew D. Halls, Front. Chem., 2022, 9, 800371
  2. Active Learning for the Design of Novel OLED Materials Hadi Abroshan, Anand Chandrasekaran, Paul Winget, Yuling An, Shaun Kwak, Christopher T. Brown, Tsuguo Morisato, and Mathew D. Halls, SID Symposium Digest of Technical Papers, 2022, 53, 885-888

Software and services to meet your organizational needs

Industry-Leading Software Platform

Deploy digital drug discovery workflows using a comprehensive and user-friendly platform for molecular modeling, design, and collaboration.

Research Enablement Services

Leverage Schrödinger’s team of expert computational scientists to advance your projects through key stages in the drug discovery process.

Scientific and Technical Support

Access expert support, educational materials, and training resources designed for both novice and experienced users.

Learning to Taste: Application of Deep Learning to Predict the Sweetness of Small Organic Molecules

AUG 6, 2020

Learning to taste: Application of deep learning to predict the sweetness of small organic molecules

Speaker: 
Atif Afzal, Senior Scientist

Abstract:
With the recent developments in machine learning techniques, we use the data available on the taste of existing chemistries to develop efficient data-driven models for taste prediction. We demonstrate that these advanced machine learning models are highly efficient in classifying the molecules as bitter/sweet. Using the data and the model developed in this work, we can not only predict the sweetness of new molecules but also identify underlying relationships that distinguish the molecules as bitter/sweet.

Accelerating materials innovation with Bunsen, your agentic co-scientist for physics & AI

Webinar

Accelerating materials innovation with Bunsen, your agentic co-scientist for physics & AI

CalendarDate & Time
  • October 6th, 2026
  • 8:00 AM PDT | 11:00 AM EDT | 4:00 PM BST | 5:00 PM CEST
LocationLocation
  • Virtual
Register

Materials R&D is often slowed not by a lack of ideas, but by the effort required to turn a scientific objective into a rigorous, executable workflow – from assembling trustworthy data and selecting appropriate methods to managing calculations, interpreting results, and deciding what to do next. Schrödinger’s Bunsen is an agentic co-scientist that connects these steps in a truly chemistry-native environment – grounded in 3D atomistic structures rather than text alone, and built on Schrödinger’s physics-based modeling, machine learning, and established scientific best practices.

In this webinar, practical materials development case studies will show how Bunsen moves beyond a conventional chat interface to plan and execute multi-step workflows spanning literature and data analysis, quantum mechanics and molecular dynamics simulations, machine learning, iterative candidate refinement, and the synthesis of results into clear, shareable reports. Across catalysis, semiconductor processing, organic electronics, energy storage, consumer-product formulations, and sustainable polymers, Bunsen helps teams navigate complex, multidimensional design spaces, identify data gaps and modeling risks, and prioritize the most informative candidates for further computation or experiment.

Attendees will see how agentic automation that is guided by scientific oversight can reduce manual handoffs, translate data into interpretable knowledge and sharable reports, preserve traceability, and help materials R&D teams move more efficiently from question to evidence-backed decision.

Our Speaker

Anand Chandrasekaran

Product Manager, AI/ML for Materials Science, Schrödinger

Anand Chandrasekaran joined Schrödinger in 2019 and he is currently the Product Manager of AI/ML for Materials Science. His expertise is in applying machine learning to different areas in Materials Science and computational modeling. He graduated from the group of Prof.Nicola Marzari in the Swiss Federal Institute of Technology, Lausanne with a PhD in Materials Science. Before joining Schrödinger, Anand also worked in the group of Prof. Rampi Ramprasad on a number of topics including polymer informatics, machine-learning force-fields, and machine-learning for electronic structure calculations.

Register

CECAM Workshop 2026

Conference

CECAM Workshop 2026

CalendarDate & Time
  • October 14th-16th, 2026
LocationLocation
  • Lyon, France

Schrödinger is thrilled to serve on the organizing committee for the upcoming workshop, “Coarse-Graining in the Age of Machine Learning and Molecular Design” on October 14th – 16th in Lyon, France.

At the event, our team will deliver a talk, “Bridging Scales in Formulation Science: Coarse-Grained Simulations with the Schrödinger Software Suite,” showcasing how tailored CG methodologies (like DPD and Martini) accelerate product design.

Don’t miss the chance to share your work at the intersection of CG modeling and machine learning. Register or submit your abstract! Discuss your work challenges with Schrödinger scientists at the workshop.

icon time
Bridging Scales in Formulation Science: Coarse-Grained Simulations with the Schrödinger Software Suite

Speaker:
Irene Bechis, Principal Scientist I, Materials Science Applications Science

Abstract:
Coarse-grained (CG) modeling has emerged as a vital approach for probing the behavior of complex molecular systems at length- and timescales beyond the reach of conventional atomistic molecular dynamics (MD). Driven by the necessity to bridge the gap between microscopic interactions and macroscopic phenomena, the field has recently experienced rapid acceleration in both foundational method development and diverse industrial applications. To keep pace with these modern advancements, the Schrödinger software suite has continuously evolved, integrating cutting-edge CG workflows and rigorously validating them against increasingly complex and challenging systems.

This presentation showcases the value of the CG approach through case studies spanning applications in the areas of pharmaceutical formulations and consumer packaged goods. Key examples will include modeling the self-assembly, mRNA encapsulation, and delivery mechanisms of lipid nanoparticles (LNPs) and simulating how complex commercial formulations interact with, penetrate, or modify biological substrates such as hair and skin. Utilizing a range of distinct CG methodologies—specifically Dissipative Particle Dynamics (DPD) and the Martini force field framework—we demonstrate how tailored simulation workflows can accelerate product design.

Collectively, these case studies illustrate how modern CG modeling within the Schrödinger ecosystem provides critical insights that enable the rational design and optimization of next-generation therapeutic and consumer products.

Physics-driven ML to accelerate the design of layered multicomponent electronic devices

FEB 10, 2026

Physics-driven ML to accelerate the design of layered multicomponent electronic devices

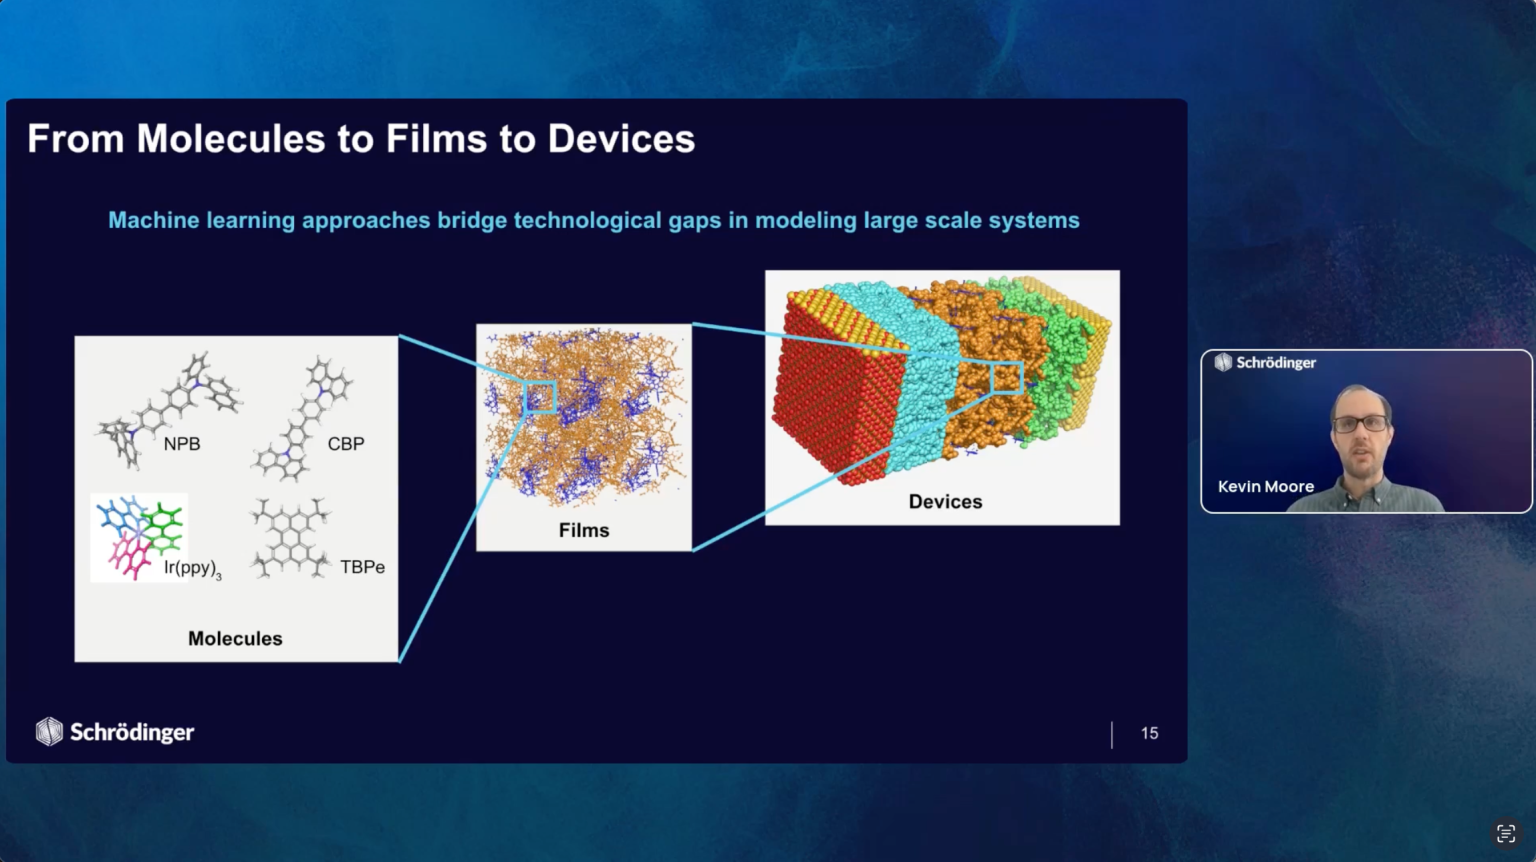
Many advanced electronic devices – such as OLEDs, batteries, solar cells, and transistors – rely on complex multilayer architectures composed of multiple materials. Optimizing device performance, stability, and efficiency requires precise control over layer composition and arrangement, yet experimental exploration of new designs is costly and time-intensive. Although physics-based simulations offer insight into individual materials, they are often impractical for full device architectures due to computational expense and methodological limitations.

Schrödinger has developed a machine learning (ML) framework that enables users to predict key performance metrics of multilayered electronic devices from simple, intuitive descriptions of their architecture and operating conditions. This approach integrates automated ML workflows with physics-based simulations in the Schrödinger Materials Science suite, leveraging physics-based simulation outputs to improve model accuracy and predictive power. This advancement provides a scalable solution for rapidly exploring novel device design spaces – enabling targeted evaluations such as modifying layer composition, adding or removing layers, and adjusting layer dimensions or morphology. Users can efficiently predict device performance and uncover interpretable relationships between functionality, layer architecture, and materials chemistry. While this webinar focuses on single-unit and tandem OLEDs, the approach is readily adaptable to a wide range of electronic devices.

Key highlights:

  • A machine learning framework for modeling electronic device performance, allowing users to define architectural features to explore novel device configurations
  • Model accuracy demonstrated with a dataset of over 2,000 OLED architectures for multiple key performance metrics
  • Pre-trained ML models for six device performance metrics available out-of-the box, including external quantum efficiency, current efficiency, power efficiency, electroluminescence maximum peak position, bandwidth, and emission color
  • Intuitive graphical interface for designing, training, and exploring new chemistries and device architectures
  • Demonstration of the framework’s extensibility to a broad range of electronic devices

Who should attend:

  • Device developers
  • R&D leaders
  • Innovation managers
  • Digitization managers
  • Synthetic chemists
  • Computational materials scientists

Our Speaker

Kevin Moore

Senior Scientist II, Materials Science Software, Schrödinger

Kevin Moore is a scientist at Schrӧdinger working on development of multiscale and hybrid physics-AI predictive frameworks for discovery and optimization of next generation materials, devices and fabrication. Prior to joining Schrӧdinger, he earned a Ph.D. in computational chemistry from the University of Georgia and conducted postdoctoral research at Argonne National Laboratory. He specializes in quantum physics-based calculations to predict the structure, properties, and reactivity of chemical systems. Recently, his efforts have been on training and validating new ML architectures and models, leveraging first-principles data, informatics and physics featurization. These new AI frameworks bridge chemistries and length scales, spanning from atoms to devices. One particular target area involves the design of electronic devices such as OLEDs, batteries, solar cells, transistors, and more.

Virtual Screening Online Course FAQ

Background

FAQ

Virtual Screening Online Course

I have registered with a start date in the future

How do I change my course start date?

You can modify your course registration here, and choose a new time frame that works for you. This is also a great page to double-check your end-dates across courses.

I am currently taking the course

How do I access the virtual cluster (i.e. how do I access Maestro)?

Your credentials for the virtual cluster that you will use to access web-based Maestro were shared in an email with the subject line “Virtual screening with integrated physics and machine learning — Welcome and Credentials!”. If you have not received that email please make sure to first check your Spam and Junk folders and let us know so we can find other ways to deliver the information. Your username with the Masetro virtual workstation will be a user number, for instance “user5”. To access the virtual cluster, choose the geographic region that is closest to you:

North America/South America access

Europe/Africa/Asia/Australia access

Please note that each Schrödinger Online Course has a different set of virtual cluster URLs. If you have enrolled in multiple courses please be sure to use the correct course-specific URL to log into the virtual workstation for the course you are currently working on. The virtual clusters require high-speed internet for ideal performance.

Please also remember that uploading or downloading files or data to web-based Maestro that are not related to the course is strictly prohibited.

Can I have an extension?

The timing of this course has been found to work well with the schedule of a participant with a full-time role. However, we understand that schedules are dynamic and we would like to ensure all participants feel they have adequate time to complete the course modules and final assignment. Due to the nature of the computational resources that are provisioned for the course, extensions are evaluated on a case-by-case basis. Please fill out this here to request an extension.

I have completed the course

How do I access my course certificate and badge?

Your certificate and badge will be available within the course platform when you reach 100% completion for your course. Be sure to complete any unfinished lessons and be aware that assignments need to be approved by a Schrödinger scientist before an assignment lesson is considered complete.

After the course, can I continue to use Maestro for my own research?

Unfortunately, the virtual cluster is only used for our online certification courses. If you are interested in using Maestro for your own research, please reach out and we can provide you more information on how to bring Schrödinger tools to your research group.