All Schrödinger offices worldwide will be closed for the week of August 17-21 as part of a company-wide initiative to rest and recharge. Please expect limited responses during this time. Scientific and Technical Support team members will be available to answer emergency support issues only.

EUROPIN Summer School on Drug Design 2025

EUROPIN Summer School on Drug Design 2025

CalendarDate & Time
  • September 14th-19th, 2025
LocationLocation
  • Vienna, Austria

Schrödinger is excited to be participating in the EUROPIN Summer School on Drug Design 2025 conference taking place on September 14th – 19th in Vienna, Austria. Join us for a presentation and workshops by Daniel Cappel, Senior Principal Scientist at Schrödinger.

icon time SEPT 16 | 14:00
A novel workflow for the in silico identification and prioritization of potential allosteric binding sites based on mixed solvent simulations and SiteMap

Speaker:
Daniel Cappel, Senior Principal Scientist, Schrödinger

Abstract:
Allosteric modulation is a promising strategy for developing drugs against difficult targets where traditional orthosteric site targeting faces challenges with selectivity, resistance, or developability. The increasing availability of high-resolution protein structures and advancements in computational power and in silico algorithms have expanded the potential of structure-based drug design (SBDD) for allosteric drug discovery. However, there remains a significant need for effective tools to identify and prioritize potential allosteric binding sites, particularly those not accessible in the apo protein structure. To address this, we have developed a novel workflow that leverages mixed solvent molecular dynamics (MxMD) simulations to reveal potential binding sites, coupled with an improved SiteMap for scoring the druggability of these sites. Our combined approach achieved a >80% top 5 found rate of known allosteric binding sites in apo structures from a set of 22 apo/holo PDBs, compared to only 63% with SiteMap alone and 54% with MxMD alone. We further evaluated our workflow on a curated set of five pharmaceutically relevant targets with multiple known allosteric binding sites, and our method outperformed popular machine learning methods, p2rank and DiffDock, as well as SiteMap, in all systems except one. Finally, we report the successful application of this workflow to an active drug discovery project within our therapeutics group. Our new workflow offers an improved method for characterizing binding sites in allosteric systems. Furthermore, efforts are underway to refine the predicted binding sites to enable virtual screening campaigns. This combined approach demonstrates a significant improvement over existing methods for identifying allosteric binding sites and has the potential to enable effective hit discovery campaigns for novel binding sites.

icon time SEPT 16 | 14:30
A beginner’s guide to system preparation, docking and designing ligands

Speaker:
Daniel Cappel, Senior Principal Scientist, Schrödinger

Abstract:
If you are interested in learning to navigate the Schrödinger suite and how to perform docking of small molecules, join us for a hands-on workshop designed for beginners. The main Maestro interface houses all the tools that are required to bring in your starting small molecules and protein system, so that they may be prepared correctly. Once you are comfortable with the fundamentals of preparing your raw materials, we will move on to understanding more about the binding site and its features, which will help us think about how a ligand might interact with it. This forms a fundamental basis for understanding our docking results, so we will start by setting up and running docking jobs and analyzing how the resulting docked compounds fulfill the basic criteria of shape and molecular interactions that lead to the final scoring term. Finally, we will explore ligand design in a more automated fashion using the Ligand Designer GUI which facilitates on-the-fly ideation through ‘build and dock’ workflows. Using embedded libraries of building-blocks, users can modify their initial idea in many intuitive ways: from attachment points on the bound ligand; the free and viable space in the binding site; through picking specific residues in the protein or specific waters in the binding cavity to guide the design process.

icon time SEPT 17 | 14:30
Efficient virtual screening: Combining ligand-based screening with QuickShape and advanced water-based scoring with WaterMap and GlideWS

Speaker:
Daniel Cappel, Senior Principal Scientist, Schrödinger

Abstract:
In this workshop, we will assemble a modern virtual screening pipeline from start to finish. We will curate a tailored screening library by simultaneously querying catalogs of many vendors. Using QuickShape screening, we will efficiently prioritize compounds for molecular docking, reducing computational cost and focusing on promising candidates. Then, we will perform compound selection leveraging chemical properties, pharmacophore features, and diversity analysis with the Hit Analyzer tool in Maestro to identify a diverse set of potential hits. Using GlideWS, we will rescore these compounds and use the advanced visualizer in Maestro to eliminate likely false positives by analyzing protein desolvation, improving the accuracy of your screening results. Finally, we will nominate a selection of top-ranking compounds for sophisticated AB-FEP+ rescoring.
Festival of Biologics Basel 2025

Conference

Festival of Biologics 2025

CalendarDate & Time
  • September 30th – October 2nd, 2025
LocationLocation
  • Basel, Switzerland

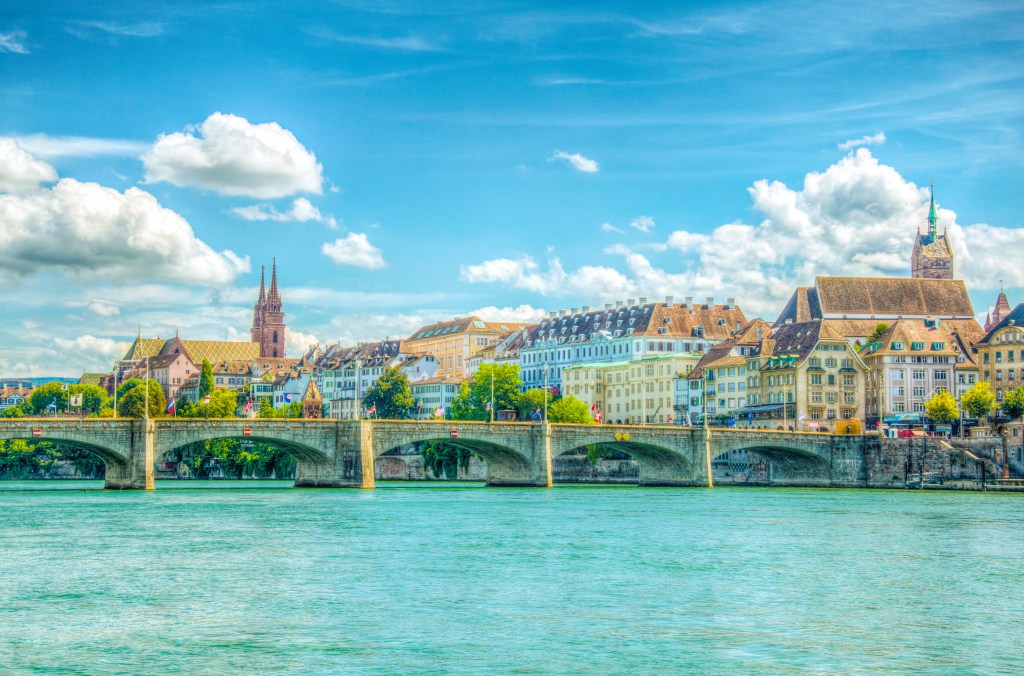
Schrödinger is excited to be participating in the Festival of Biologics 2025 conference taking place on September 30th – October 2nd in Basel, Switzerland. Join us for a presentation by Esam Abualrous, Principal Scientist I at Schrödinger, titled “Advances in Structure-Based Computational Modeling and Collaborative Enterprise Informatics for Biologics.” Stop by booth 501B to speak with Schrödinger scientists.

icon time OCT 1 | 14:40 CET
icon location Theatre 6
Advances in Structure-Based Computational Modeling and Collaborative Enterprise Informatics for Biologics

Speaker:
Esam Abualrous, Principal Scientist I, Schrödinger

Abstract:
Optimizing biologic drug candidates to enhance favorable traits or minimize liabilities often demands significant experimental effort. This presentation will showcase recent progress in our physics-based, structure-guided computational methods that help accelerate the optimization process. It will cover key challenges in antibody engineering—including predicting antibody-antigen binding affinity, improving structural stability, and addressing developability concerns. This talk will illustrate how these approaches can be integrated within a collaborative informatics platform for biologics discovery, which can merge machine learning (ML) results, experimental data and advanced modeling, execution, and analysis tools to streamline decision-making and centralize essential project information.

Release 2025-3

Library Background

Release Notes

Release 2025-3

Small Molecule Drug Discovery

Platform Environment

Maestro Graphical Interface

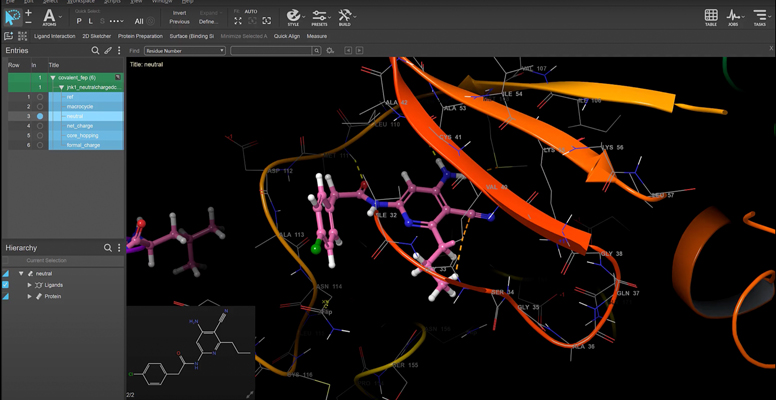
  • Hovering the cursor over an atom in the Workspace now simultaneously highlights its corresponding row in the Project Table and displays its row number in the Status Bar
  • Improved defaults to “Get PDB Dialog” to auto display 2Fo-Fc diffraction data and auto set isocontours in EM maps to the Author recommendation
  • Redesigned 2D Viewer Export with dedicated options dialog to control image size and support for high-quality SVG format for both single structures and HTML grids
  • New Maestro Assistant (beta) – An AI-powered conversational interface providing context-aware help via the Schrödinger Knowledge Bot and enabling natural language styling command execution within the 3D workspace (documentation)

Force Field

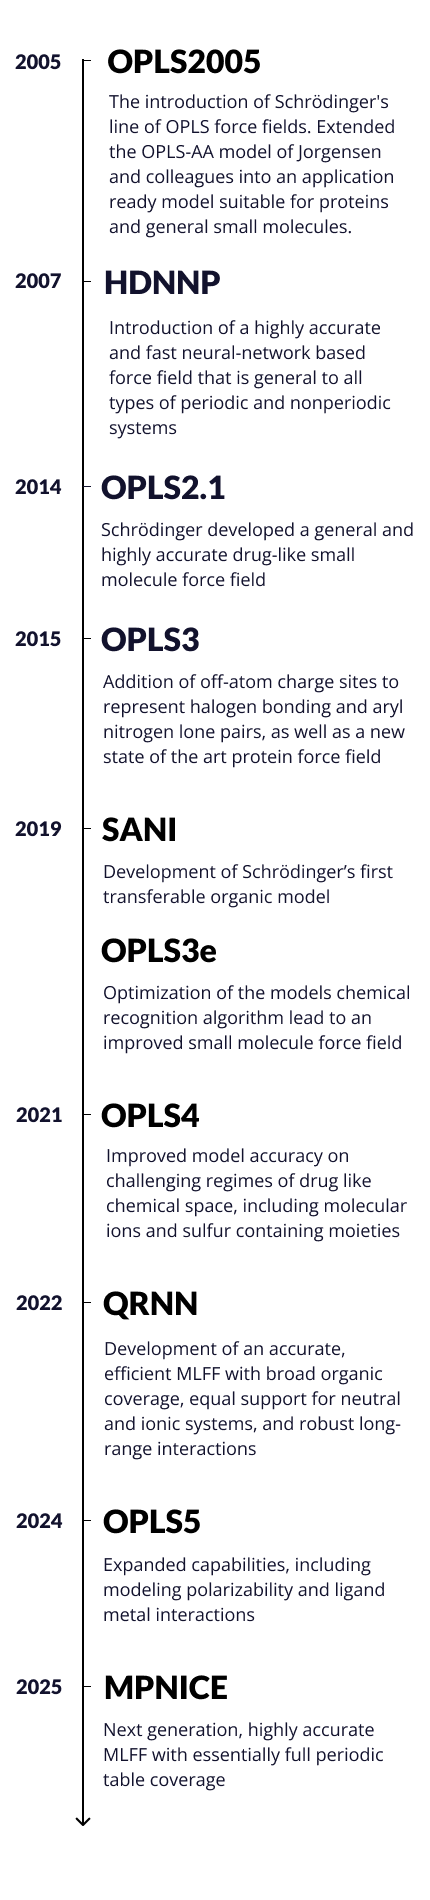
  • Updated FFBuilder default reference method to the MPNICE potential
    • Roughly 5x faster for typical chemistries
    • 20x faster for boron, bromine, iodine and silicon containing compounds

Target Validation & Structure Enablement

Protein Preparation

  • Protein Preparation Wizard interactive jobs now save temporary files under a named folder in the Maestro working directory

Protein X-Ray Refinement

  • Improved defaults to “Get PDB Dialog” to auto display 2Fo-Fc diffraction data and auto set isocontours in EM maps to the Author recommendation
  • Redesigned GlideMap panel to dock ligands into maps generated from X-ray data by GlideXtal

Cryo-EM Model Refinement

  • Redesigned GlideMap panel replaces the GlideEM panel with more granular options for fitting small molecule ligands into cryo-EM density maps

IFD-MD

  • New Template Ligand Finder for ligand-binding mode prediction and cryptic binding site identification (Beta): Rapidly identify and visualize homologous proteins based on both protein and ligand similarity to uncover potential template ligands as references for IFD-MD

Binding Site & Structure Analysis

Mixed Solvent MD (MxMD)

  • Enhanced cryptic binding site identification with updated in silico workflow (Beta): New workflow combining mixed solvent molecular dynamics (MxMD) with SiteMap to reliably reveal and detect cryptic binding sites

Hit Identification & Virtual Screening

Docking

  • Optimized Glide which is nearly 2x faster than Glide is now the default method
  • Revamp of the Glide WS MMGBSA correction

Ligand Preparation

Macrocycles

  • Add command-line options to restrain cis/trans isomerism during alignment with tug_align.py
  • Add a command-line option to tug_align.py to control MCS search algorithm timeout
  • Small performance improvements to PrimeMCS

ABFEP

  • Automatic membrane placement for AB-FEP simulations

Lead Optimization

FEP+

  • New Settings interface that replaces the Advanced Options interface for more intuitive and easy simulation set up
  • Clear predictions for RB-FEP edges
  • Similarity Score column displayed in the analysis tab
  • Batch delete and download multiple jobs with enhanced usability of the ‘Web Services Jobs Table’ panel
  • Extend Atom Mapping to matched R-groups

Constant pH Simulations

  • Improved panel usability and layout
  • Added support to run constant pH simulations via Web Services

FEP Protocol Builder

  • Roughly 2X speedup in workflow through improved defaults
  • Improved accuracy in generated FEP+ maps through exploration and scoring of submaps with Louvian clustering. Alternatively, users can input desired submaps
  • New cost-optimal option using Pareto Selection of FEP-PB Models to generate the best value protocols
  • Improved prediction accuracy of generated maps through FEP+ Groups support, where ligands with different protonation, tautomeric, and conformational forms will be grouped, enabling FEP Group corrections to be applied

Quantum Mechanics

  • Predict Ames toxicity via a QM-based workflow following Leach et al. (2009)
  • X-ray emission spectroscopy (XES) prediction is now available (command line only)
  • Implemented nine new double hybrid functionals with RI-MP2: B2-PLYP, B2GP-PLYP, DSD-BLYP, DSD-PBEP86, PWPB95, B2K-PLYP, B2T-PLYP, DSD-PBEB95, MPW2-PLYP
  • Wave function stability analysis automatically corrects SCF instabilities leading to a more stable wave function
  • Predict Nucleus-Independent Chemical Shifts (NICS) with new workflow
  • Optical rotation as a function of wavelength
  • Employ MLFFs (machine learning force fields) by setting Level of Theory option
  • MPNICE now supported in all Jaguar workflows that have supported use of QRNN

Life Science Education Content

Biologics Drug Discovery

  • Easily specify mutational variants to be modeled in MMGBSA Residue Scanning by uploading an input FASTA file
  • Load, analyze and visualize in the MSV desired mutations specified in the MMGBSA Residue Scanning Panel by newly available export of variants to a FASTA file
  • Macromolecular Pose Filtering supports multi-chain ligands when filtering based on data obtained from Hydrogen-Deuterium Exchange (HDX) experiments
  • Added a predefined selection for the Vα-Vβ interface, located under the TCR Regions menu, to enable one-click selection of this key binding region
  • New T cell receptor (TCR)-specific presets

Materials Science

GUI for Quantum ESPRESSO

Product: Quantum ESPRESSO (QE) Interface

  • Defect Correction: (+DEFECT_FORMATION_ENERGY) Formation energy computed in the panel
  • Finite displacement method for phonons (command line)
  • Increased number of iterations and cycles for default setup
  • Support for GBRV pseudopotentials by default

MS Surface

Product: MS SurfChem

  • Desorption Enumeration: (+ASSOCIATIVE_DESORPTION) Option for associative desorption

Microkinetics

Product: MS Microkinetics

  • Calculation of selectivity from Catalytic Reaction Analysis (command line)

Reactivity

Product: MS Reactivity

  • Reaction Network Profiler: Support for Garza solvation entropy partition functions (command line)
  • Nanoreactor: Improvements to elementary reaction network algorithms

KMC Charge Mobility

Product: MS Mobility

  • Option to compute field-dependent mobility based on charge diffusion (command line)

Dielectric properties

Product: MS Dielectric

  • Complex Permittivity: Option to apply custom fitting parameters
  • Complex Permittivity: Display of predictions from multiple fits
  • Complex Permittivity: Option to input a pre-equilibrated structure
  • Complex Permittivity: Improved speed from KWW parameter calculations

Reactive Interface Simulator

Product: MS RIS

  • Solid Electrolyte Interphase: Support for custom reactions that modify bond orders (command line)
  • Solid Electrolyte Interphase: Improved algorithm to track unpaired electrons

Crystal Structure Prediction

Product: Crystal Structure Prediction

  • Crystal Structure Prediction: Simplified UI for improved UX
  • Crystal Structure Prediction: All space groups shown in Advanced Settings

Advanced Force Field Applications

Product: MS FF Applications

  • Machine learning force field support in QM and MD panels

Transport Calculations via MD simulations

Product: MS Transport

  • Diffusion: Support for GPU calculations on driver host
  • Ionic Conductivity: (+IONIC_CONDUCTIVITY) Workflow solution to predict ionic conductivity in liquids
  • Thin Plane Shear: (+PLANE_SHEAR_VELOCITY_PROFILE) Reduced noise in velocity profile
  • Thin Plane Shear: (+PLANE_SHEAR_VELOCITY_PROFILE) Direct calculation of viscosity velocity profile
  • Thin Plane Shear: Improved control of shear area to avoid drift of slabs

Coarse-Grained (CG) Molecular Dynamics

Product: MS CG

  • Speed up for assignment of the coarse-grained force fields for large systems
  • Coarse-grained Mapping: GUI panel for automatic mapping of CG structures
  • CG FF Builder: Automated addition of antifreeze water for MARTINI mapping
  • CG FF Builder: (+CGFF_BUILDER_MARTINI_NPT) Support for NPT ensemble during the fitting of MARTINI force field parameters
  • CG FF Builder: Support for encrypted force field parameters
  • CG FF Builder: Support for loading output from CG Mapping as input

Materials Informatics

Product: MS Informatics

  • Machine Learning Property: Tooltip to visualize the training set chemical space
  • MD Descriptors: Automated setup for label and number of components from input CSV

Formulation ML

Product: MS Formulation ML

  • Support for the description of ingredients without SMILES strings
  • Formulation ML: Visualization of model performance from the GUI
  • Formulation ML: Option to export feature importance to CSV
  • Formulation ML: Option to compute feature importance during training
  • ML Model Manager: GUI for management of model and version information

Layered Device ML

Product: MS Layered Device ML

  • OLED Device ML: Support for classification models

MS Maestro Builders and Tools

  • Adsorption Enumeration: Improved adsorption for sterically hindered atoms
  • Adsorption Enumeration: De-duplication for molecular adsorption
  • Complex Builder: Title suggestions for sketched ligands based on IUPAC name
  • Complex Builder: Build dimeric organometallic complexes
  • Disordered System: Speed-up for snap-to-grid and amorphous modes
  • Disordered System: Support for MLFF
  • Meta Workflows: Support for MLFF
  • Polymer: Setup for angles on coarse-grained polymer models

Classical Mechanics

  • Complex Bilayer: (+COMPLEX_BILAYER_BUILDER) Model building solution for complex protein membrane systems
  • Diffusion: Support for MLFF
  • Polymer Crosslink: Speed up with improved crosslinking algorithms
  • Electrolyte Analysis: Option to plot density distribution isosurface
  • MD Multistage: Preset relaxation protocol for stiff polymers
  • MD Multistage: Preset relaxation protocol to aid convergence in OPLS5
  • MD Multistage: Option to choose electric field units
  • MD Multistage: Option to view steps from the preset relaxation protocols
  • MD Multistage: Support for MLFF
  • Molecular Deposition: Support for MLFF
  • Polymer Chain Analysis: Improved speed on searching for backbone atoms
  • Prepare for MD: Support for MLFF
  • Radial Distribution Function panel restored for user access
  • Visualize Restraints: Tool to show restraints in systems for MD simulations

Quantum Mechanics

  • Adsorption Energy: Support for MLFF
  • Beta Elimination: Support for MLFF
  • Bond and Ligand Dissociation: Molecular formulas printed for fragments
  • Nanoreactor: Support for MLFF
  • Optoelectronic Film Properties: Option to calculate the reorganization energies for ISC/RISC
  • Optoelectronic Film Properties: Option to plot the distribution of singlet-triplet splittings for ISC/RISC in the viewer panel
  • Probe Grid Scan: Support for MLFF
  • Reaction Energetics Enumeration: Support for MLFF
  • Reaction Network Profiler: Support for MLFF

Materials Science Education Content

Education Content

Life Science

Materials Science

LiveDesign

What’s New in 2025-3

  • Freeform Columns
    • The Freeform column user interface to define picklist values has been redesigned to facilitate faster picklist value creation
    • Upload 3D attachments to a Freeform column (‘pse’, ‘mae’, ‘maegz’, ‘mmtf’, ‘pdb’, ‘pdbgz’, ‘sdf’) and view them in the 3D Visualizer
  • Plots
    • Pinned tooltips are included in plot exports
    • Box plots now permit horizontal and vertical overlays
  • Advanced Search
    • “Case-insensitive” and “Match any” options are now available as gear options in Advanced search queries
    • The “Real” and “Virtual” quick filter options are now available for all entity types in the Advanced search panel
    • Scrolling is now enabled while running auto-update advanced search. Pointer actions for individual search conditions and the top of the drawer are disabled
  • Sequence Viewer
    • View the chemical structure for non-natural residues with docked tooltip
    • Residues that match the corresponding reference residue can be represented as dots(.) instead of letters, to more easily identify differences
    • Drag selection now extends during click and drag until the end of the canvas
    • Data columns section in the sequence viewer now auto opens if we toggle the data columns checkbox in the collapsed state
  • Data & Columns Tree
    • Published limited assay columns are now searchable via the [LIM] prefix in the D&C tree and typing the complete column name searches and highlights the respective column
  • Import
    • Admin users can define any existing user as ther “Lot Scientist” for an entity using LDClient with the following requirements: 1) the username must exactly match an existing username in LiveDesign; new usernames cannot be created and other strings cannot be used, and 2) the Lot Scientist can only be defined when registering a new entity, and cannot be updated for existing entities
  • Admin Panel
    • Admin users can update the ENABLE_TECHNICAL_USERS property to show Technical users within user lists
    • Bulk kill up to 40,000 tasks in Task Engine
  • UX Improvements
    • The one-click “View in Maestro” link has been redesigned to more clearly separate the “View in 3D” and “View in Maestro” options
    • Sorting icons on the column headers have been updated
    • Columns that are filtered in the LiveReport now show a filter icon on the column header
    • The confirmation dialog for copying compounds from one LiveReport to another has been removed
    • Admin users can configure the number of entities that can be viewed simultaneously in the 3D visualizer by defining the MAX_LOADED_3D_STRUCTURES property
    • Column headers remain visible when scrolling within the Stereoisomer tool

What’s Been Fixed

  • Admin Panel
    • License files would occasionally fail to upload in the Admin Panel UI, and now correctly upload
    • Models would show hyper-restricted projects as an optional project to add the Model to, even though the Protocol did not include the hyper-restricted project. The Models page now only shows the projects that are included in the Protocol
  • Freeform columns
    • Picklist Freeform columns would show incorrect coloring rules if a picklist value was selected that contained a newline, and now show the correct colors
    • Creating a Comment Freeform column would fail if the column name matched the title of any other column in any other LiveReport, and now succeeds, unless the column name matches an existing published Freeform column title
  • Filters
    • Toggling off filters on a LiveReport would show flashing cells until the browser was refreshed, and now shows the correct cell values’
  • Forms
    • Copying text from a widget in Forms would fail if the widget was configured as a drilldown widget, and now text can be copied from drilldown widgets
    • Characters would occasionally get deleted while typing in the Forms annotation widget, and now do not get deleted
  • Formulas
    • Formulas returning multiple values would not show values when a if() function was used, and now correctly shows multiple values
  • Ligand Designer
    • Ligand Designer: reference ligands using a .maegz format would not show up in the 3D visualizer, and now correctly show up in the 3D Visualizer
  • Limited Assay Columns
    • Pasting strings into the Create Limited Assay Column dialog would briefly appears, but then disappear, and now remains visible in the dialog
  • LiveReports
    • Clicking a LiveDesign hyperlink would fail to open the linked LiveReport, and instead would open the user’s last-opened LiveReport; clicking hyperlinks now correctly opens the linked LiveReport
    • Copying a LiveReport from one project to another would fail when the LiveReport contained a “Presence in LiveReport” Advanced Search condition, and now succeeds.
  • Templates
    • Applying a template to a LiveReport would not show the applied filters unless the browser page was refreshed, and now correctly shows the applied filters
  • The “Give Feedback” button has been disabled

Training & Resources

Online Certification Courses

Level up your skill set with hands-on, online molecular modeling courses. These self-paced courses cover a range of scientific topics and include access to Schrödinger software and support.

Tutorials

Learn how to deploy the technology and best practices of Schrödinger software for your project success. Find training resources, tutorials, quick start guides, videos, and more.

Other Resources

Force Field Bundle

Force Field Bundle

Improve the quality of your computational predictions with best-in-class, proprietary force fields — developed in-house and built for accuracy

Force Field Bundle

Updated Force Field Bundle offers you expanded coverage into more complex systems

We’re advancing force field innovations to help you achieve more accurate, reliable modeling outcomes

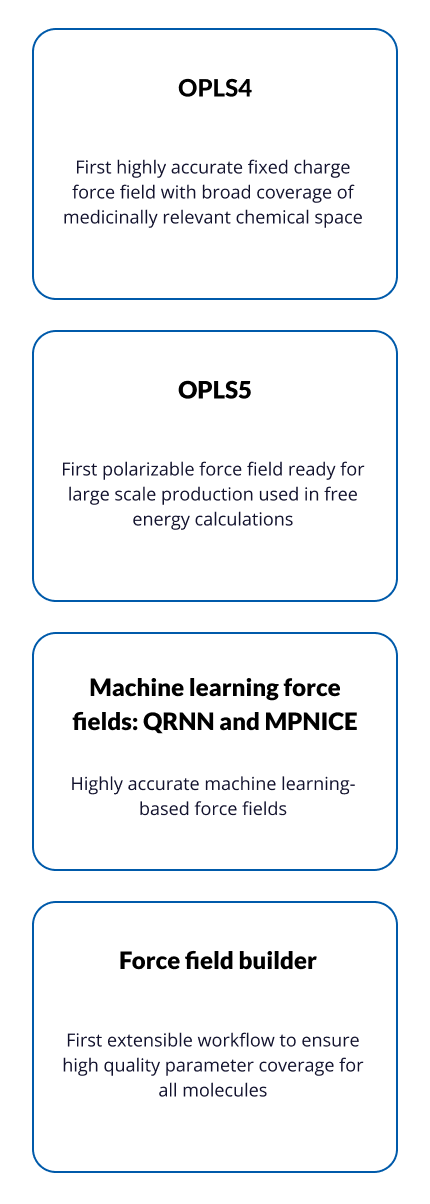
Force fields are used in molecular simulations to describe the interactions between atoms in a system. Having an accurate force field is at the heart of obtaining useful molecular structures and predicting relative energies, and yet many in silico programs employ force fields that are years, if not decades, old and suffer from lack of sufficient coverage for many common molecular motifs.

Introducing OPLS5: Enabling Innovation in Molecular Design With an Advanced Force Field

We are witnessing a shift toward a computational “predict-first” approach to drug discovery and materials science research.

Key Benefits of Schrödinger Force Fields

  • Continuous scientific development by leading force field experts
  • Backed by a state of the art quantum engine (Jaguar) and extensive experimental validation
  • Broad coverage of chemical space for small molecules, biologics and materials science applications
  • Easily extendible into novel project-specific chemistry with the Force Field Builder
  • Highly accurate and scalable machine learning force fields with broad expandable coverage and reach of system

Applications for drug discovery

Obtain more accurate predictions of binding affinity

Generate precise binding free energy predictions with FEP+, enabling more reliable rank ordering within congeneric series.

Expand the domain of applicability with machine learning force fields

Enhance and accelerate physics-based computational methods by integrating AI/ML into force fields and simulation engines.

Predict binding modes of novel scaffolds

Accurately predict binding modes of novel scaffolds using advanced induced fit docking methods in IFD-MD.

Perform accurate molecular dynamics simulations

Reveal mechanisms of action and key interaction energies through high-fidelity molecular dynamics simulations with Desmond.

Improve conformational analyses

Achieve better conformational sampling and docking poses with improved torsional energy descriptions across Glide, ConfGen, MacroModel, and Prime.

Related Products

FEP+

High-performance free energy calculations for drug discovery

IFD-MD

Accurate ligand binding mode prediction for novel chemical matter, for on-targets and off-targets

Desmond

High-performance molecular dynamics (MD) engine providing high scalability, throughput, and scientific accuracy

Force Field Builder

Efficient tool for optimizing custom torsion parameters in OPLS4

MS Force Field Applications

Cutting-edge force field technologies for accurate property predictions

Related Publications

Life Science Publication

A robust crystal structure prediction method to support small molecule drug development with large scale validation and blind study

Life Science Publication

Towards automated physics-based absolute drug residence time predictions

Life Science Publication

Accurate physics-based prediction of binding affinities of RNA- and DNA-targeting ligands

Materials Science Publication

Gaining molecular insights towards inhibition of foodborne fungi Aspergillus fumigatus by a food colourant violacein via computational approach

Materials Science Publication

Predicting Drug-Polymer Compatibility in Amorphous Solid Dispersions by MD Simulation: On the Trap of Solvation Free Energie

Materials Science Publication

Possible Applications of the Polli Dissolution Mechanism: A Case Study Using Molecular Dynamics Simulation of Bupivacaine

Materials Science Publication

Modelling of Prednisolone Drug Encapsulation in Poly Lactic-co-Glycolic Acid Polymer Carrier Using Molecular Dynamics Simulations

Materials Science Publication

Cu-TiO2/Zeolite/PMMA Tablets for Efficient Dye Removal: A Study of Photocatalytic Water Purification

Life Science Publication

Coarse-grained simulation of mRNA-loaded lipid nanoparticle self-assembly

Life Science Publication

OPLS5: Addition of polarizability and improved treatment of metals

Training & Resources

Online certification courses

Level up your skill set with hands-on, online molecular modeling courses. These self-paced courses cover a range of scientific topics and include access to Schrödinger software and support.

Tutorials

Learn how to deploy the technology and best practices of Schrödinger software for your project success. Find training resources, tutorials, quick start guides, videos, and more.

XXIX National Meeting on Medicinal Chemistry

Conference

XXIX National Meeting on Medicinal Chemistry

CalendarDate & Time
  • September 14th-17th, 2025
LocationLocation
  • Parma, Italy

Schrödinger is excited to be participating in the XXIX National Meeting on Medicinal Chemistry conference taking place on September 14th – 17th in Parma, Italy. Join us for a presentation by Giulia D’Arrigo, Senior Scientis I at Schrödinger, titled “Accelerating Drug Discovery through Integrated Physics-Based and Machine Learning Approaches using Schrödinger’s Computational Platform.”

Speaker:

Giulia D’Arrigo, Senior Scientis I, Schrödinger

Abstract:

Schrödinger’s computational platform accelerates molecular design and drug discovery by integrating physics-based methods and machine learning to address the complexities of multi-parameter optimization using a comprehensive suite of in silico tools and workflows spamming the entire research and development pipeline. Here, real-world applications of these technologies will be illustrated across different case studies from the Schrödinger Therapeutics Group (STG).

Ultra-large scale chemical space exploration using AutoDesigner [1,2] allows the generation and efficient prioritization of potentially billions of novel structures to systematically explore novel chemical scaffolds or R-groups that meet project requirements. Combining machine learning with rigorous free energy calculations using FEP+ [3,4] in an iterative fashion (Active Learning FEP+), further allows to rapidly score the ultra-large libraries generated by AutoDesigner to identify designs with optimal potency and selectivity profile. Examples of this automated large scale approach are the identification, after 7 months of project initiation, of novel WEE1 inhibitors with >10,000X selectivity over PLK [1] as well as the discovery of four novel scaffolds of EFGR inhibitors in only six days [5].

A new, in-silico approach for addressing ADME/T liabilities and enabling precise control over key endpoints is presented. This approach leverages state of the art computational tools such as induced-fit docking (IFD-MD) [6,7], FEP+, and solvation energy calculations with E-sol [8], in order to enable a rational approach to ADME. Applications of these workflows to hERG inhibition, CYP rate of metabolism, and brain exposure and efflux are presented. . A prime example is the in silico-enabled discovery of KAI-11101 [9], a DLK inhibitor for neurodegenerative disease treatment. Here, accurate ADMET profiling and large scale on-target and off-target FEP+ were fundamental to elect KAI-11101 as a brain penetrant, potent and highly selective kinase inhibitor preclinical candidate.

Altogether, these examples demonstrate the impact of Schrödinger’s computational platform in facilitating and accelerating drug discovery programs.

References

[1] Bos et al. J Chem Inf Model. 2024 Oct 14;64(19):7513-7524.
[2] Bos et al. J. Chem. Inf. Model. 2022, 62, 8, 1905–1915.
[3] Wang et al.  J. Am. Chem. Soc., 2015, 137(7), 2695–2703.
[4] Ross et al. Commun. Chem., 2023, 6(222).
[5] Igawa et al. J Med Chem. 2024 Dec 26;67(24):21811-21840.
[6] Miller EB, et al. Cell, 2024, 187, 3, 521-525.
[7] Miller et al. J. Chem. Theory Comput. 2021, 17, 4, 2630–2639.
[8] Lawrenz et al. J Chem Inf Model. 2023 Jun 26;63(12):3786-3798.
[9]Lagiakos et al. J Med Chem. 2025 Feb 13;68(3):2720-2741.

MS Force Field Applications

MS Force Field Applications

Cutting-edge force field technologies for accurate property predictions

MS Force Field Applications

MS Force Field Applications (MS FF Applications) includes access to Schrödinger’s widely-used OPLS4 and new OPLS5 force field, as well as all the Schrödinger machine learning force fields (MLFFs) for diverse property prediction workflows. Schrödinger is committed to advancing innovations in force fields to help you achieve more accurate, reliable modeling outcomes.

What’s New: 

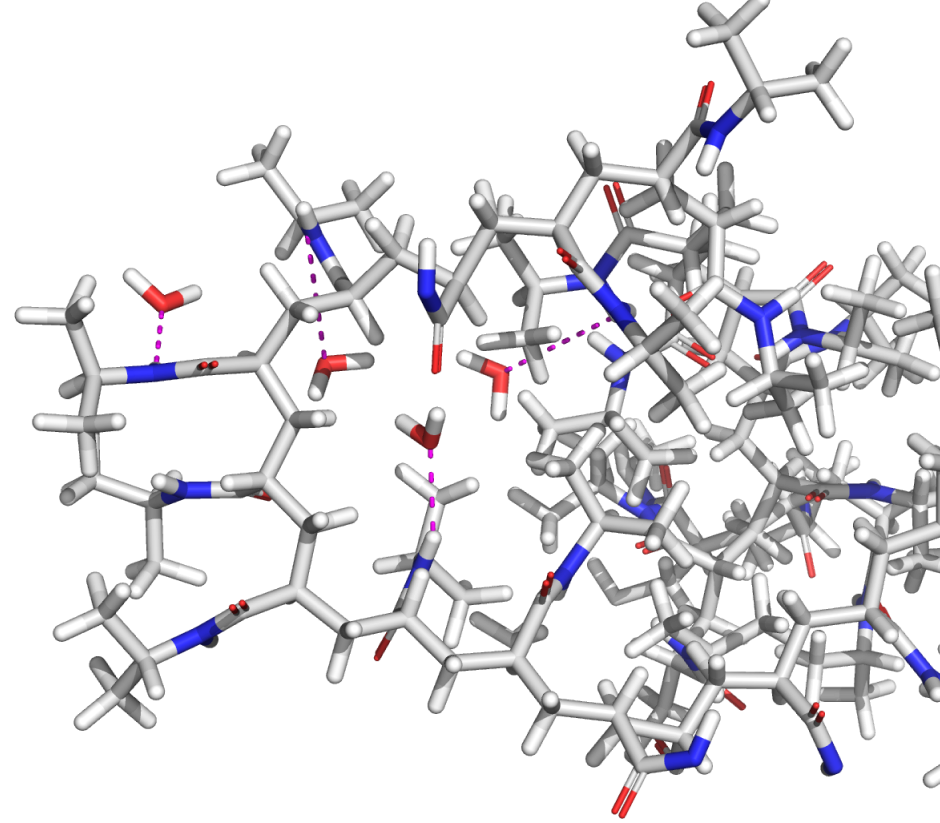
  • OPLS5: Includes an explicit treatment of polarizability via the addition of Drude oscillators that enables accurate modeling of cation-pi interactions and more accurate treatment of hydrogen bonding to charged systems.
  • MPNICE: Machine learning force fields, also known as machine learning interatomic potentials, represent an intermediate between classical force fields and DFT, maintaining the linear scaling of the former while approaching the accuracy of the latter. Message Passing Network with Iterative Charge Equilibration (MPNICE) is an MLFF architecture developed by Schrödinger for which multiple pre-trained models covering 89 elements are available, and which explicitly incorporates equilibrated atomic charges and long range electrostatics.
  • Fine-tuning: While architectures like MPNICE and UMA offer exceptional generalizability, achieving high-fidelity accuracy for specialized systems often requires tailoring the model through quantum mechanical data. MS FF Applications provides an intuitive, automated interface for fine-tuning MPNICE models using DFT calculations from Jaguar or Quantum Espresso. This enables the development of custom, application-specific MLFFs that integrate seamlessly across the full range of materials science workflows and solutions.
  • MPNICE embeddings for 3D ML models: The internal layers of MLFFs encapsulate a sophisticated, dense mapping of chemical space and 3D atomic configurations. By leveraging these latent representations, MPNICE embeddings allow you to build machine learning models directly from 3D coordinates. This enables the creation of high-fidelity predictive models across organic, inorganic, and hybrid systems – spanning both periodic and non-periodic structures.
background pattern

Benefits of MLFFs

Near DFT-level accuracy with orders of magnitude reduction in computational time
Option for GPU accelerated molecular dynamics with Desmond
Large chemical space spanning 89 elements
Specialized force fields for organics, inorganics, and hybrid materials with the added capability of fine-tuning

Key Capabilities

Batteries

  • Calculate bulk and transport properties, such as diffusion, viscosity, and conductivity, of liquid electrolytes
  • Simulate Li-ion diffusion in solid state electrolytes and cathode coating materials
  • Model electrolyte reactivity and SEI formation

Polymers

  • Evaluate polymer dynamical properties
  • Investigate solid polymer electrolyte

Adsorption on surfaces

  •  Study reactivity of multiple adsorbates in extended models of complex surfaces

Crystal structure prediction

  • Rank order organic crystal structures

OLED materials

  • Simulate molecular packing and thin-film morphology 
  • Investigate doping, host–guest, and interlayer interactions
  • Link device properties to the static and dynamic disorder of molecular systems
  • Facilitate thermomechanical property prediction
  • Model charge and exciton transport

Case studies & webinars

Discover how Schrödinger technology is being used to solve real-world research challenges.

Materials Science Webinar

Advancing battery materials innovation using charge-aware machine learning force fields

In this webinar, we will demonstrate how Schrödinger is utilizing an integrated computational approach combining physics-based molecular modeling with machine learning force fields (MLFFs) to address key challenges in battery materials design.

Materials Science Webinar

Advancing machine learning force fields for materials science applications 最新機能 MPNICEのご紹介

シュレーディンガーが開発した最先端のMLFFアーキテクチャ「MPNICE(Message Passing Network with Iterative Charge Equilibration)」をご紹介します。

Materials Science Webinar

Advancing machine learning force fields for materials science applications

In this webinar, we will introduce Schrödinger’s state-of-the-art MLFF architecture, called Message Passing Network with Iterative Charge Equilibration (MPNICE), which incorporates explicit electrostatics for accurate charge representations.

Documentation & Tutorials

Get answers to common questions and learn best practices for using Schrödinger’s software.

Materials Science Tutorial

Machine Learning with MPNICE Embedding

Learn to build, analyze, and apply ML models with MPNICE Embedding.

Materials Science Tutorial

Fine-Tuning Machine Learning Force Fields

Learn to fine-tune a pre-trained machine learning force field (MLFF) model to study lattice parameters of 2D materials.

Materials Science Tutorial

Locating Adsorption Sites on Surfaces

Learn how to locate adsorption sites on surfaces.

Materials Science Documentation

Machine Learning Force Fields

Machine Learning Force Fields (MLFFs) offer a novel approach for predicting the energies of arbitrary systems.

Materials Science Quick Reference Sheet

MLFF Calculations: Quick Reference Sheet

Get an overview of the MLFF Calculations panel for predicting quantum mechanical calculations for systems using machine learning force fields.

Materials Science Tutorial

Machine Learning Force Field

Learn how to use machine learning force field optimization methods to prepare and simulate various systems.

Related Products

OPLS4

Modern, comprehensive force field for accurate molecular simulations

Desmond

High-performance molecular dynamics (MD) engine providing high scalability, throughput, and scientific accuracy

Force Field Builder

Efficient tool for optimizing custom torsion parameters in OPLS4

MS Maestro

Complete modeling environment for your materials discovery

Jaguar

Quantum mechanics solution for rapid and accurate prediction of molecular structures and properties

Quantum ESPRESSO Interface

Integrated graphical user interface for nanoscale quantum mechanical simulations

Publications

Materials Science Publication

Efficient long-range machine learning force fields for liquid and materials properties

Materials Science Publication

Leveraging high-throughput molecular simulations and machine learning for the design of chemical mixtures

Materials Science Publication

Advancing material property prediction: using physics-informed machine learning models for viscosity

Materials Science Publication

Machine learning force field ranking of candidate solid electrolyte interphase structures in Li-ion batteries

Schedule a demo on MS Force Field Applications

Contact us today to discuss how you can leverage MLFFs to solve your R&D challenges.

Don’t see your areas of interest in the current lists above? Reach out so we can help.

Form submitted

Thank you, we’ll be in touch soon.

Software & services to meet your organizational needs

Software Platform

Deploy digital materials discovery workflows with a comprehensive and user-friendly platform grounded in physics-based molecular modeling, machine learning, and team collaboration.

Research Services

Leverage Schrödinger’s expert computational scientists to assist at key stages in your materials discovery and development process.

Support & Training

Access expert support, educational materials, and training resources designed for both novice and experienced users.

IMID 2025

Conference

IMID 2025

CalendarDate & Time
  • August 19th-22nd, 2025
LocationLocation
  • Busan, Korea

Schrödinger is excited to be participating in the 25th International Meeting on Information Display conference taking place on August 19th – 22nd in Busan, Korea. Mathew D. Halls, Senior Vice President of Materials at Schrödinger, will be chairing the session “AI for Efficient Display Design” on Wednesday, Aug 20.

Join us for a presentation by Mathew D. Halls, titled “From Molecules to Displays: A Digital Chemistry Platform Uniting Physics-Based Simulation with Machine Learning for Optoelectronic Design”. Stop by our booth to speak with Schrödinger scientists.

icon time AUG 20 | 10:50AM
icon location Room F (313)
From Molecules to Displays: A Digital Chemistry Platform Uniting Physics-Based Simulation with Machine Learning for Optoelectronic Design

Speaker:
Mathew D. Halls, Senior Vice President of Materials, Schrödinger

Abstract:
Experimental exploration of innovative architectures and material compositions for OLED devices requires substantial time, labor, and resources, due to the complexity and cost of device fabrication, characterization, and analysis. Predictive modeling offers a powerful alternative, enabling efficient and targeted evaluation of devices across broad design spaces by integrating informatics and large-scale property predictions. In this presentation, Schrödinger will showcase recent advances of its digital platform combining machine learning (ML) technologies with quantum mechanics and molecular dynamics. The first advancement extends our automated ML algorithm for chemical formulations [1] to predict performance parameters of multicomponent layered devices, e.g., OLEDs. The second advancement involves overcoming limitations of classical force fields by using a message-passing neural network potential with iterative charge equilibration to achieve quantum mechanical accuracy at minimal computational cost. These ML models encode device components (i.e., material structures, layer architectures, physicochemical properties, and operating conditions) as features to predict OLED device performance metrics for operational output, stability, and efficiency (Fig. 1). This approach moves beyond traditional chemical modeling strategies, capturing complex relationships between device architecture, composition and function. Complementing this development are advances in Schrödinger’s physics-based simulation software, which computes determinative properties of OLED materials. MPNICE, the latest version of our ML potential, delivers accurate DFT-quality predictions at reduced computational cost, enabling simulation of increasingly more complex films and processes. For example, systems combining metal and organic chemistries typically outside the coverage of traditional force fields can now be more efficiently explored. Schrödinger’s new solutions for optoelectronic materials development and device optimization provides unprecedented capabilities for accelerated development of innovative display technologies.

In silico cryptic binding site detection and prioritization

JUL 30, 2025

In silico cryptic binding site detection and prioritization

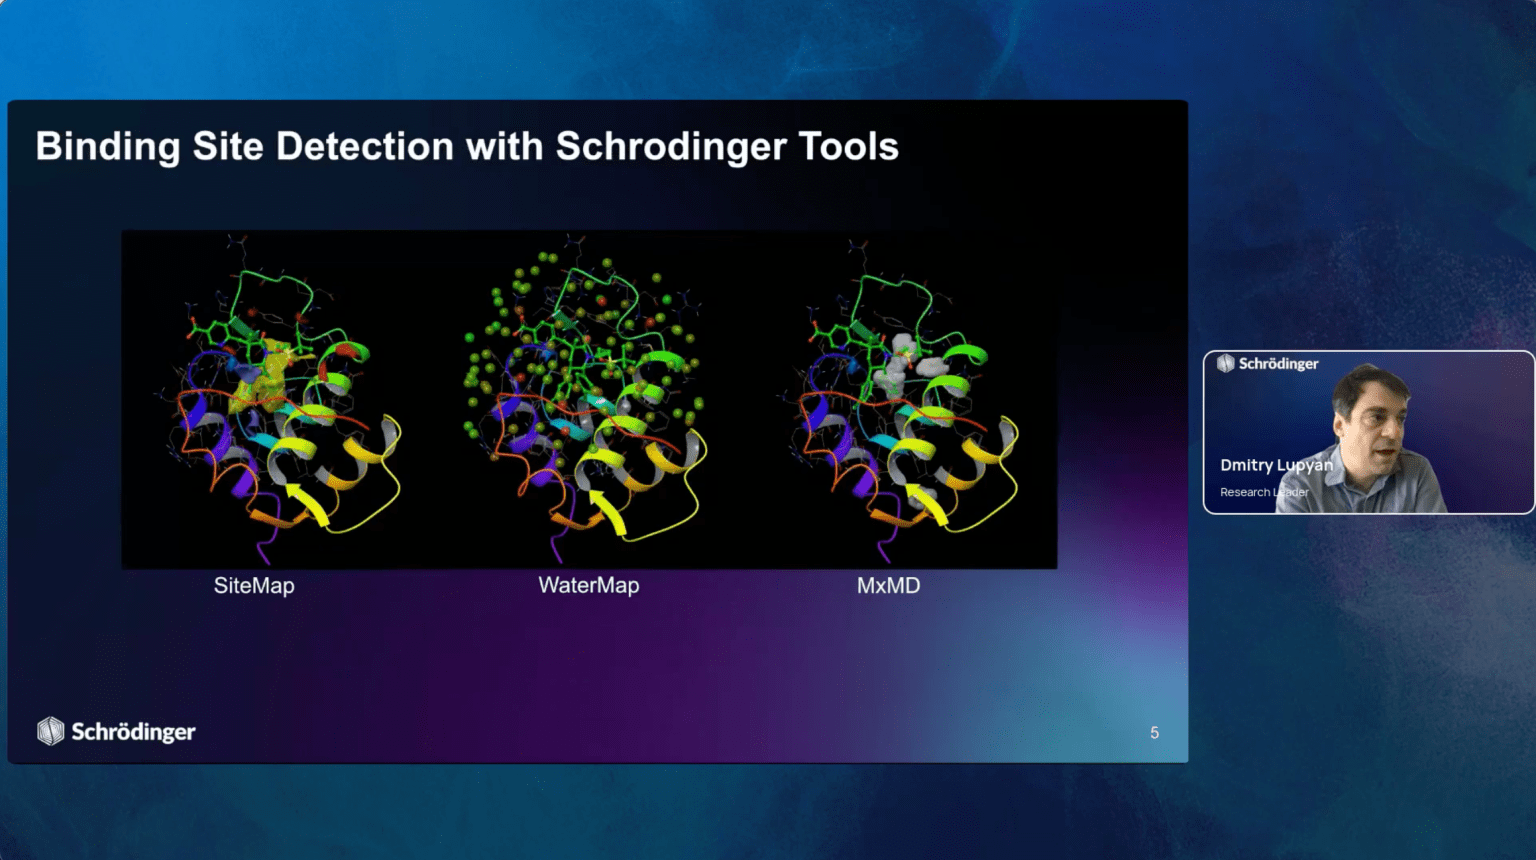
Targeting cryptic binding sites is becoming an increasingly powerful strategy for tackling challenging drug targets, especially where traditional orthosteric approaches fall short due to issues like selectivity, resistance, or poor developability. However, identifying and evaluating cryptic binding sites—especially cryptic sites not visible in apo structures—remains a key challenge in early drug discovery.

In this webinar, we will introduce a novel computational workflow that integrates mixed solvent molecular dynamics (MxMD) with SiteMap to reveal and identify cryptic binding sites. This new combined workflow achieved a remarkable 83% success rate in detecting the cryptic binding sites within a retrospective benchmark set of 61 targets.

Join us to learn how this new workflow can support the identification of cryptic binding sites and enable more structure-based drug discovery campaigns for novel targets.

Webinar Highlights

  • Overview of the MxMD method
  • Introduction of new MxMD+SiteMap workflow to identify cryptic binding sites
  • Benchmarking the new workflow against popular machine learning methods and SiteMap in its default pocket detection mode

Our Speakers

Da Shi

Principal Scientist I, Life Science Software, Schrödinger

Da Shi is a Principal Scientist in the Hit Discovery team at Schrödinger. He obtained his Ph.D. at the University of California San Diego with the supervision of Prof. Ruben Abagyan. After graduation, he worked at the Frederick National Laboratory for Cancer Research as a Data Scientist on developing machine learning platforms for drug discovery. In 2021, he joined Schrödinger and worked as an All Access Applications Scientist. He later transitioned to the Hit Discovery team working on developing workflows on cryptic binding site identification and FEP ligand pose generation.

Dima Lupyan

Senior Principal Scientist, Life Science Software, Schrödinger

Dr. Dmitry Lupyan, a product manager, spearheads the development of Desmond and FEP analysis tools, showcasing his expertise in the realm of molecular dynamics. Notably, he’s behind the Python API for simulation analysis, a cornerstone utilized across Schrödinger’s MD, MxMD, and FEP+ products. Driven by a passion for scientific advancement, he actively promotes the utilization of simulation analysis tools, fostering a community of exploration. His research interests delve into the intricate domains of protein engineering, membrane-bound systems, and the fascinating dynamics of unbinding kinetics.

Contract Research Services

Contract Research Services

Contract Research Services

Expert research support customized for your materials science R&D needs

Leverage Schrödinger’s scientific and engineering expertise

Apply advanced simulation tools to solve your materials science research challenges

Free up time and resources

Let our scientists execute the project while collaborating closely with your team

Data security is of utmost importance

Contract research customers retain all intellectual property

Best suited for companies and teams:

  • Who want to reduce time spent on trial-and error experiments
  • Who want to leverage Schrödinger’s advanced physics-based and machine learning methods and expertise
  • Who want to gain molecular-level insight into their materials
  • Who want to explore the benefits of digital approaches before investing in in-house adoption


Schrödinger’s industry leading technologies:

  • Quantum mechanics modeling
  • Molecular dynamics simulation
  • Molecular mechanics
  • Advanced AI/ML/Active Learning/De novo design
  • Machine learning force fields (MLFFs)

Advance your materials R&D with unrivaled technologies and expertise

Benefit from the full impact of Schrödinger technologies at scale

Services include all computing, licensing, and service hours required to perform comprehensive simulation projects tailored to your R&D needs.

Leverage flexible, customized solutions to ensure project success

No internal software, hardware, or computational resources needed. Benefit from expert knowledge transfer and training throughout and beyond the project. Get tailored tools and solutions designed specifically to meet your project goals.

Complement your domain expert knowledge with our expertise

After collaborating to scope out a work product, dedicated Schrödinger experts in both materials applications and digital simulations execute your projects.

Proven success

Customers across industries have trusted us with their research needs.

Panasonic
Bridgestone

To learn more about our service areas in detail, download the full flyer

Software and services to meet your organizational needs

Software Platform

Deploy digital materials discovery workflows with a comprehensive and user-friendly platform grounded in physics-based molecular modeling, machine learning, and team collaboration.

Research Services

Leverage Schrödinger’s expert computational scientists to assist at key stages in your materials discovery and development process.

Support & Training

Access expert support, educational materials, and training resources designed for both novice and experienced users.

How to find a druggable target: A computational perspective

JUN 25, 2025

How to find a druggable target: A computational perspective

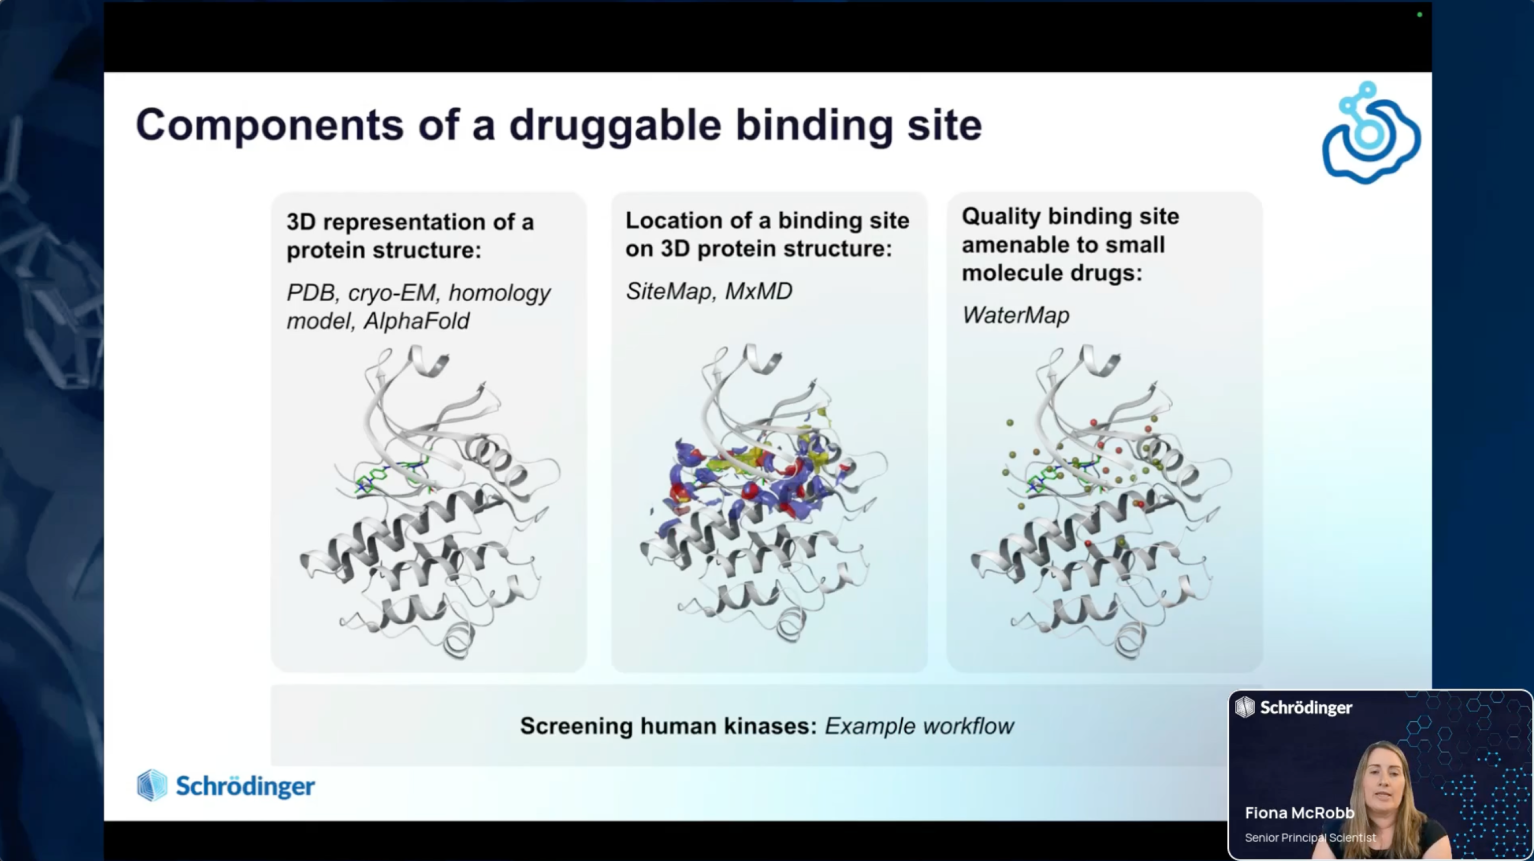
Roughly 20,000 proteins make up the human proteome, yet not all proteins are suitable drug targets. This makes finding druggable targets a critical early step in drug discovery. Structure-based techniques can enable better, quicker, and more cost effective target tractability assessment. These technologies can also expedite hit discovery and ligand binding potency and selectivity optimization and guide medicinal chemistry rationale.

Join us in this beginner-friendly webinar that will introduce you to strategies and best-in-class tools for identifying druggable, technology-enabled targets. We will review workflows to assess protein binding sites and identify protein-ligand complexes suitable for structure-based drug design. We will also discuss how to reduce uncertainty in structure-based drug models using free energy methods (FEP+), as well as examine how modern physics-based modeling approaches amplified by machine learning can reveal new therapeutic opportunities.

Last, we will highlight a real-world case study from Schrödinger’s Therapeutics Group, showcasing how these strategies were used to assess the druggability of the Wee1 binding site and subsequently how FEP+ was used to enable the discovery of a highly selective Wee1 inhibitor for the treatment of solid tumors. The case study demonstrates the power of computational approaches to unlock challenging targets and drive successful drug discovery programs.

Whether you’re new to modeling or looking to expand your toolkit, this session will provide a practical foundation and a comprehensive guide to help you get started.

Webinar highlights:

  • Discussion of what a computationally druggable target is and how to identify one
  • Overview of structure-based techniques, including docking, molecular dynamics, and free energy methods
  • Introduction to integrated tools and workflows for binding site detection, druggability assessment, and target prioritization
  • Leveraging FEP+ to validate structural models and reduce uncertainty by comparing predicted and experimental ligand affinities
  • Case study: Applying these approaches in Schrödinger’s Wee1 inhibitor discovery program

Our Speaker

Fiona McRobb

Senior Principal Scientist, Computational Chemistry, Therapeutics Group, Schrödinger

Fiona McRobb received her Ph.D. in Medicinal Chemistry from Monash University in Australia in 2012. Her Ph.D. encompassed both synthetic and computational approaches to study G protein-coupled receptors (GPCRs). In 2011, she joined Prof. Ruben Abagyan’s lab at UCSD as a postdoc, studying allosteric modulators of GPCRs and environmental toxicology. Fiona joined Schrödinger in 2014 as an Applications Scientist before transitioning to Schrödinger’s Therapeutics Group in 2016. As a Senior Principal Scientist, she applies Schrödinger’s computational chemistry software to address challenging problems in drug discovery, with a focus on identifying new targets and early drug discovery programs.

Digital and AI-Driven Materials Innovation

Symposium

Digital and AI-Driven Materials Innovation

CalendarDate & Time
  • July 8th, 2025
LocationLocation
  • Hong Kong

We hope you can join us at the Hong Kong University of Science and Technology (HKUST) on July 8th for the Digital and AI-Driven Materials Innovation symposium.

Co-organized by the Office of Knowledge Transfer and the Department of Chemistry, this premier industry-academic forum brings together leading experts, pioneering technologists, and academic innovators to explore how artificial intelligence is transforming materials science.

Attendees will:

  • Gain insights into how AI and computational tools are transforming materials discovery, from atomic-scale simulations to industrial applications
  • Learn from leading experts across academia and industry, including speakers from industry, R&D centers and academics
  • Explore real-world case studies on AI-enabled innovations in semiconductors, OLEDs, specialty polymers, and pharmaceutical formulations
  • Engage in discussions on the future of materials informatics, high-throughput screening, and machine-learned force fields
  • Network with scientists, engineers, and innovators driving the next wave of digital and AI-driven materials research

Through keynote presentations, technical talks, and a panel discussion, participants will gain actionable insights into how AI is accelerating materials innovation across sectors—from lab discovery to commercial impact.

icon time 09:30 – 10:00
Guest Registration

HKUST Clear Water Bay Campus

icon time 10:00 – 10:05
Welcome Remarks

Speaker: Prof. Tim Cheng, VPRD, HKUST

icon time 10:05 – 10:10
Opening Remarks

Speaker: Prof. Jianzhen Yu, Head and Chair Professor, Department of Chemistry, HKUST

icon time 10:10 – 10:15
Opening Remarks

Speaker: Dr. Michael Rauch, Associate Director of Materials Science, Schrödinger

icon time 10:15 – 10:20

Session Chair: Zhenyang Lin, HKUST

icon time 10:20 – 10:40
Physics-Enabled AI for Next-Generation Materials Discovery: Bridging Atomic-Scale Simulation and Machine Learning with Schrödinger

Speaker: Dr. Michael Rauch, Associate Director of Materials Science, Schrödinger

icon time 10:40 – 11:10
Materials Design in Electronics Industry: Application of Materials Informatics and High-Throughput Calculations

Speaker: Dr. Nobuyuki N. Matsuzawa, Director, Panasonic Corporation

icon time 11:10 – 11:30
Designing Tomorrow’s Materials: The Role of AI in Exchange-Correlation Functional

Speaker: Prof. Guanhua Chen, Chair Professor, Department of Chemistry, The University of Hong Kong

icon time 11:30 – 2:00
Lunch

Session Chair: Kevin Chen, HKUST

icon time 2:00 – 2:20
AI in Semiconductor Industry

Speaker: Dr. Ziyang Gao, Senior director, Hong Kong Microelectronics Research and Development Institute

icon time 2:20 – 2:40
Simulations of Atomic Level Processing with Examples from Metal ALD

Speaker: Dr. Simon Elliott, Senior Research Leader, Schrödinger

icon time 2:40 – 3:00
AI-Enabled OLED Materials Discovery

Speaker: Dr. Wei Xu, Research scientist, TCL AI Lab

icon time 3:00 – 3:20
Coffee break

Session Chair: Ping Gao, HKUST

icon time 3:20 – 3:40
AI-Guided Interfacial Engineering for Rational Design of Polymeric Microcapsules

Speaker: Prof. Jinglei Yang, Professor, Department of Mechanical and Aerospace Engineering, HKUST

icon time 3:40 – 4:00
Advancing Specialty Polymer Innovation Using Molecular Simulation and Machine Learning

Speaker: Dr. Atif Afzal, Principal Scientist, Schrödinger

icon time 4:00 – 4:20
Accelerating Pharmaceutical Formulations using the Schrödinger Software Suite

Speaker: Dr. Sudharsan Pandiyan, Principal Scientist, Schrödinger

icon time 4:20 – 5:00
Panel discussion / Closing remarks

Our Speakers

Tim Cheng

Professor, VPRD, HKUST

Jianzhen Yu

Professor, Chem, HKUST

Dr. Michael Rauch

Associate Director of Materials Science, Schrödinger

Dr. Nobuyuki N. Matsuzawa

Director, Panasonic Corporation

Dr. Ziyang Gao

Senior Director, Hong Kong Microelectronics Research and Development Institute

Dr. Wei Xu

Research Scientist, TCL AI Lab

Guanhua Chen

Professor, The University of Hong Kong

Dr. Simon Elliott

Senior Research Leader, Schrödinger

Dr. Atif Afzal

Principal Scientist, Schrödinger

Dr. Sudharsan Pandiyan

Principal Scientist, Schrödinger

Jinglei Yang

Professor, HKUST

CRS 2025

Conference

CRS 2025

CalendarDate & Time
  • July 14th-18th, 2025
LocationLocation
  • Philadelphia, Pennsylvania

Schrödinger is excited to be participating in the CRS Annual Meeting 2025 taking place on July 14th – 18th in Philadelphia, Pennsylvania. Join us for a presentation by Schrödinger’s John Shelley and David Nicholson, titled “Molecular modeling and machine learning for drug formulation.” Stop by booth #302 to speak with Schrödinger scientists.

icon time JUL 15 | 9:00AM – 10:00AM
Molecular modeling and machine learning for drug formulation

Speakers:
John Shelley, Fellow, Schrödinger
David Nicholson, Principal Scientist I, MS Applications Scientist, Schrödinger

Abstract:
Schrödinger, a leader in the application of structure-based molecular modeling and machine learning to drug discovery and development, will provide a brief, broad overview on the emerging field of structure-based molecular modeling for drug formulation followed by two more focused presentations: 1. Modeling the formation, structure and behavior of mRNA-containing lipid nanoparticles and 2. Excipient/polymer selection for small-molecule drug formulation including amorphous solid dispersions. We anticipate that attendees will gain an understanding of:
The value and limitations of structure-based molecular modelling and machine learning for drug formulation
Employing modeling to impact excipient selection
How molecular modeling, particularly when combined with experiment, can lead to a competitive advantage by providing insight into complex formulations